Decoding ferroptosis via integrated multi-omics analyses in rice blast
- 07 October 2026
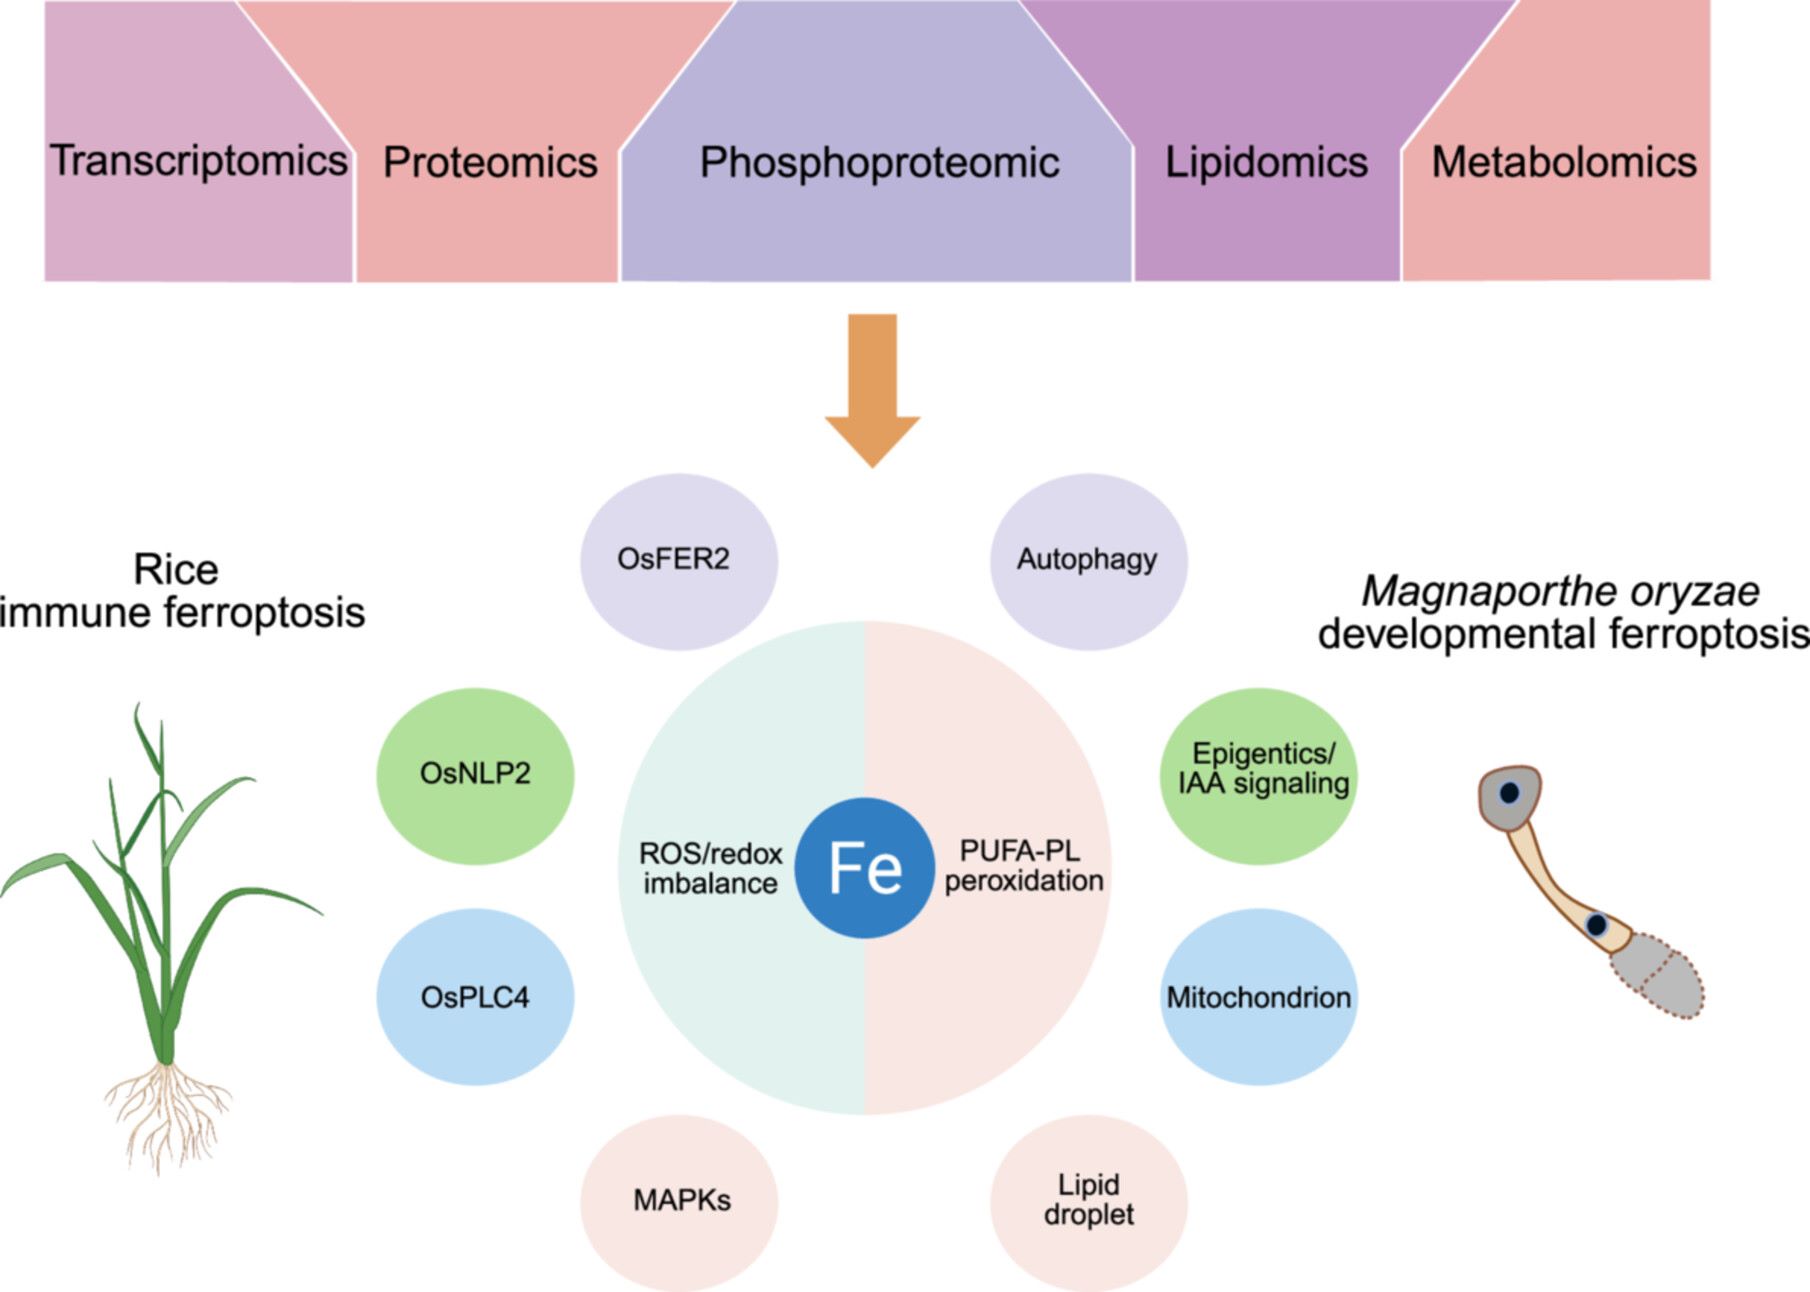
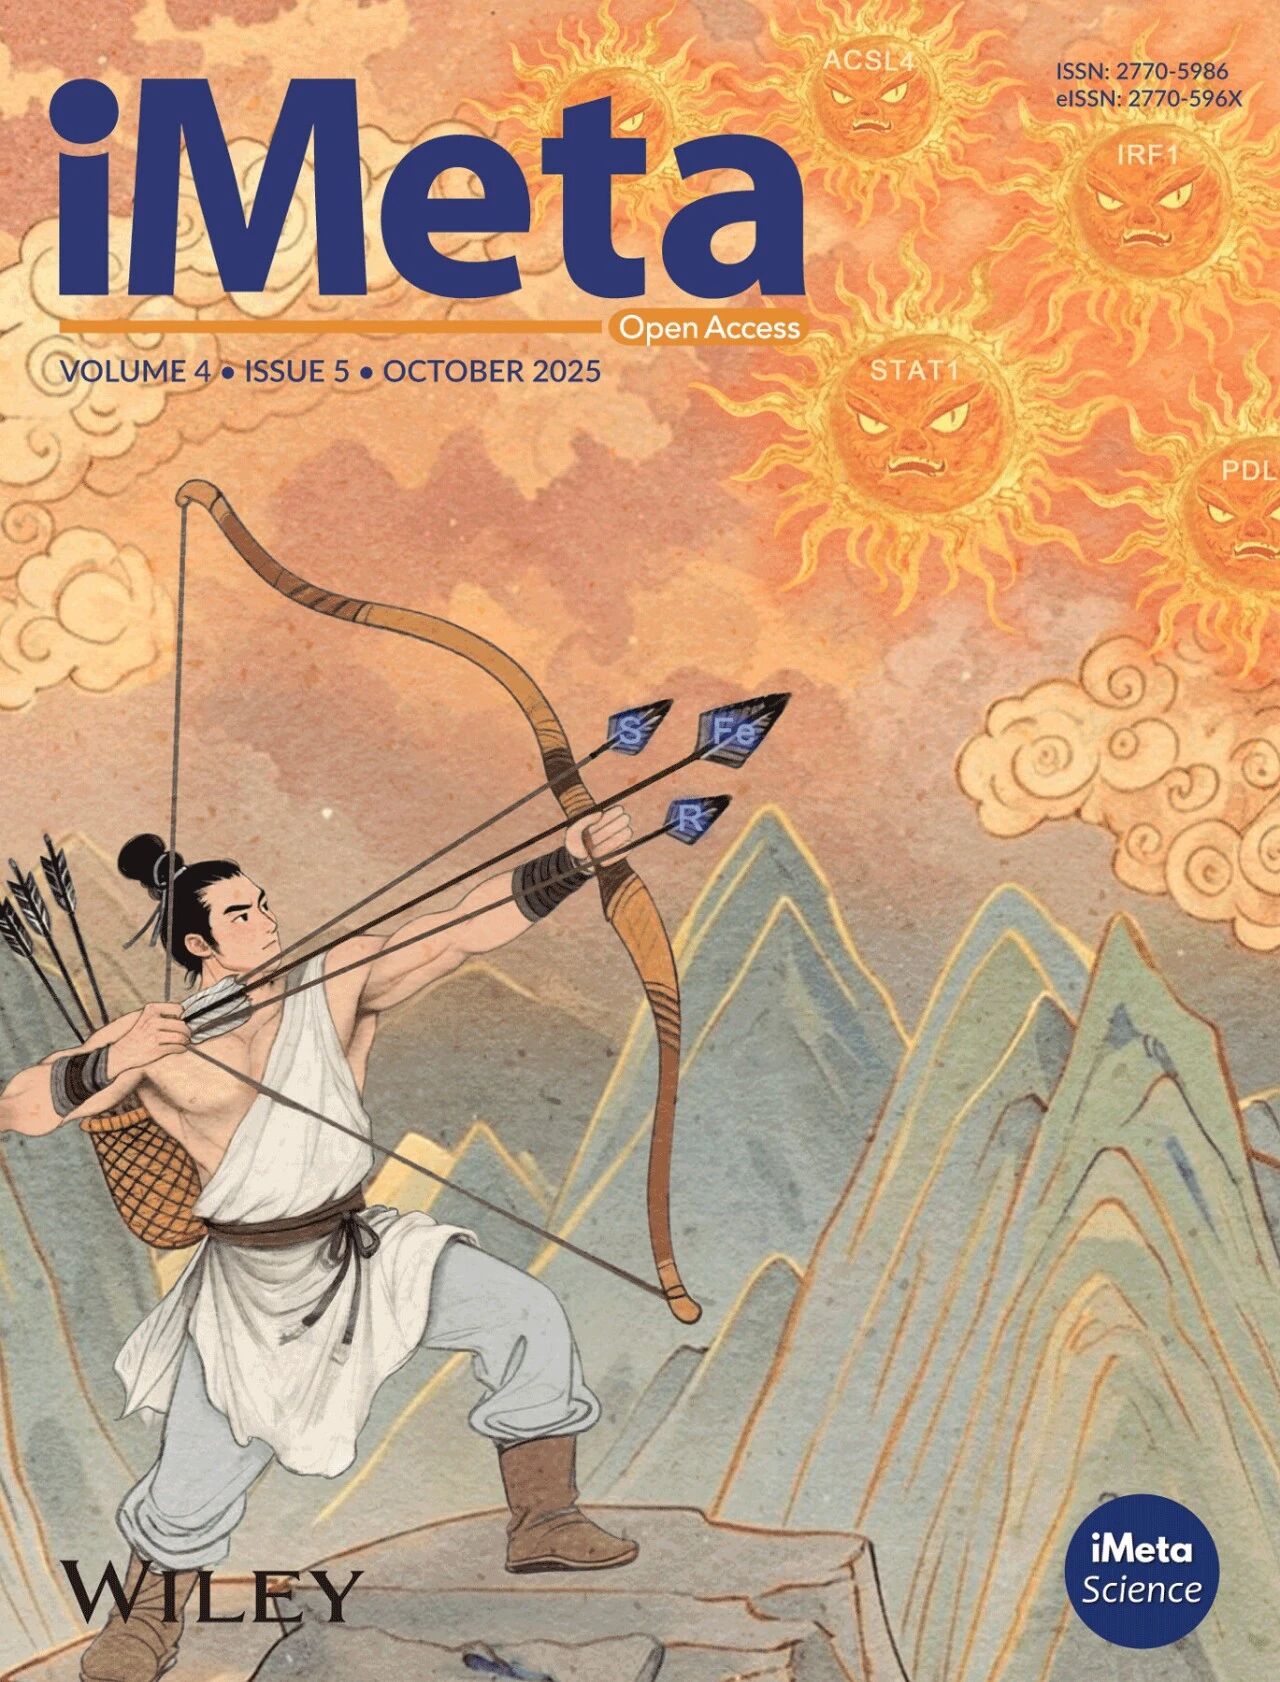
The rice-Magnaporthe oryzae pathosystem is presented as a host-pathogen ferroptosis model. Multi-omics insights connect ferroptosis to several cellular processes, such as iron homeostasis, reactive oxygen species (ROS)/redox imbalance, and polyunsaturated fatty acid-containing phospholipid (PUFA-PL) peroxidation, under multiple regulators in rice and M. oryzae, respectively. Interestingly, ferroptosis drives two opposite biological outcomes: fungal infection-related development and rice immunity.
Gut virome orchestrates nitrogen partitioning between the host and microbiome
- 04 October 2026
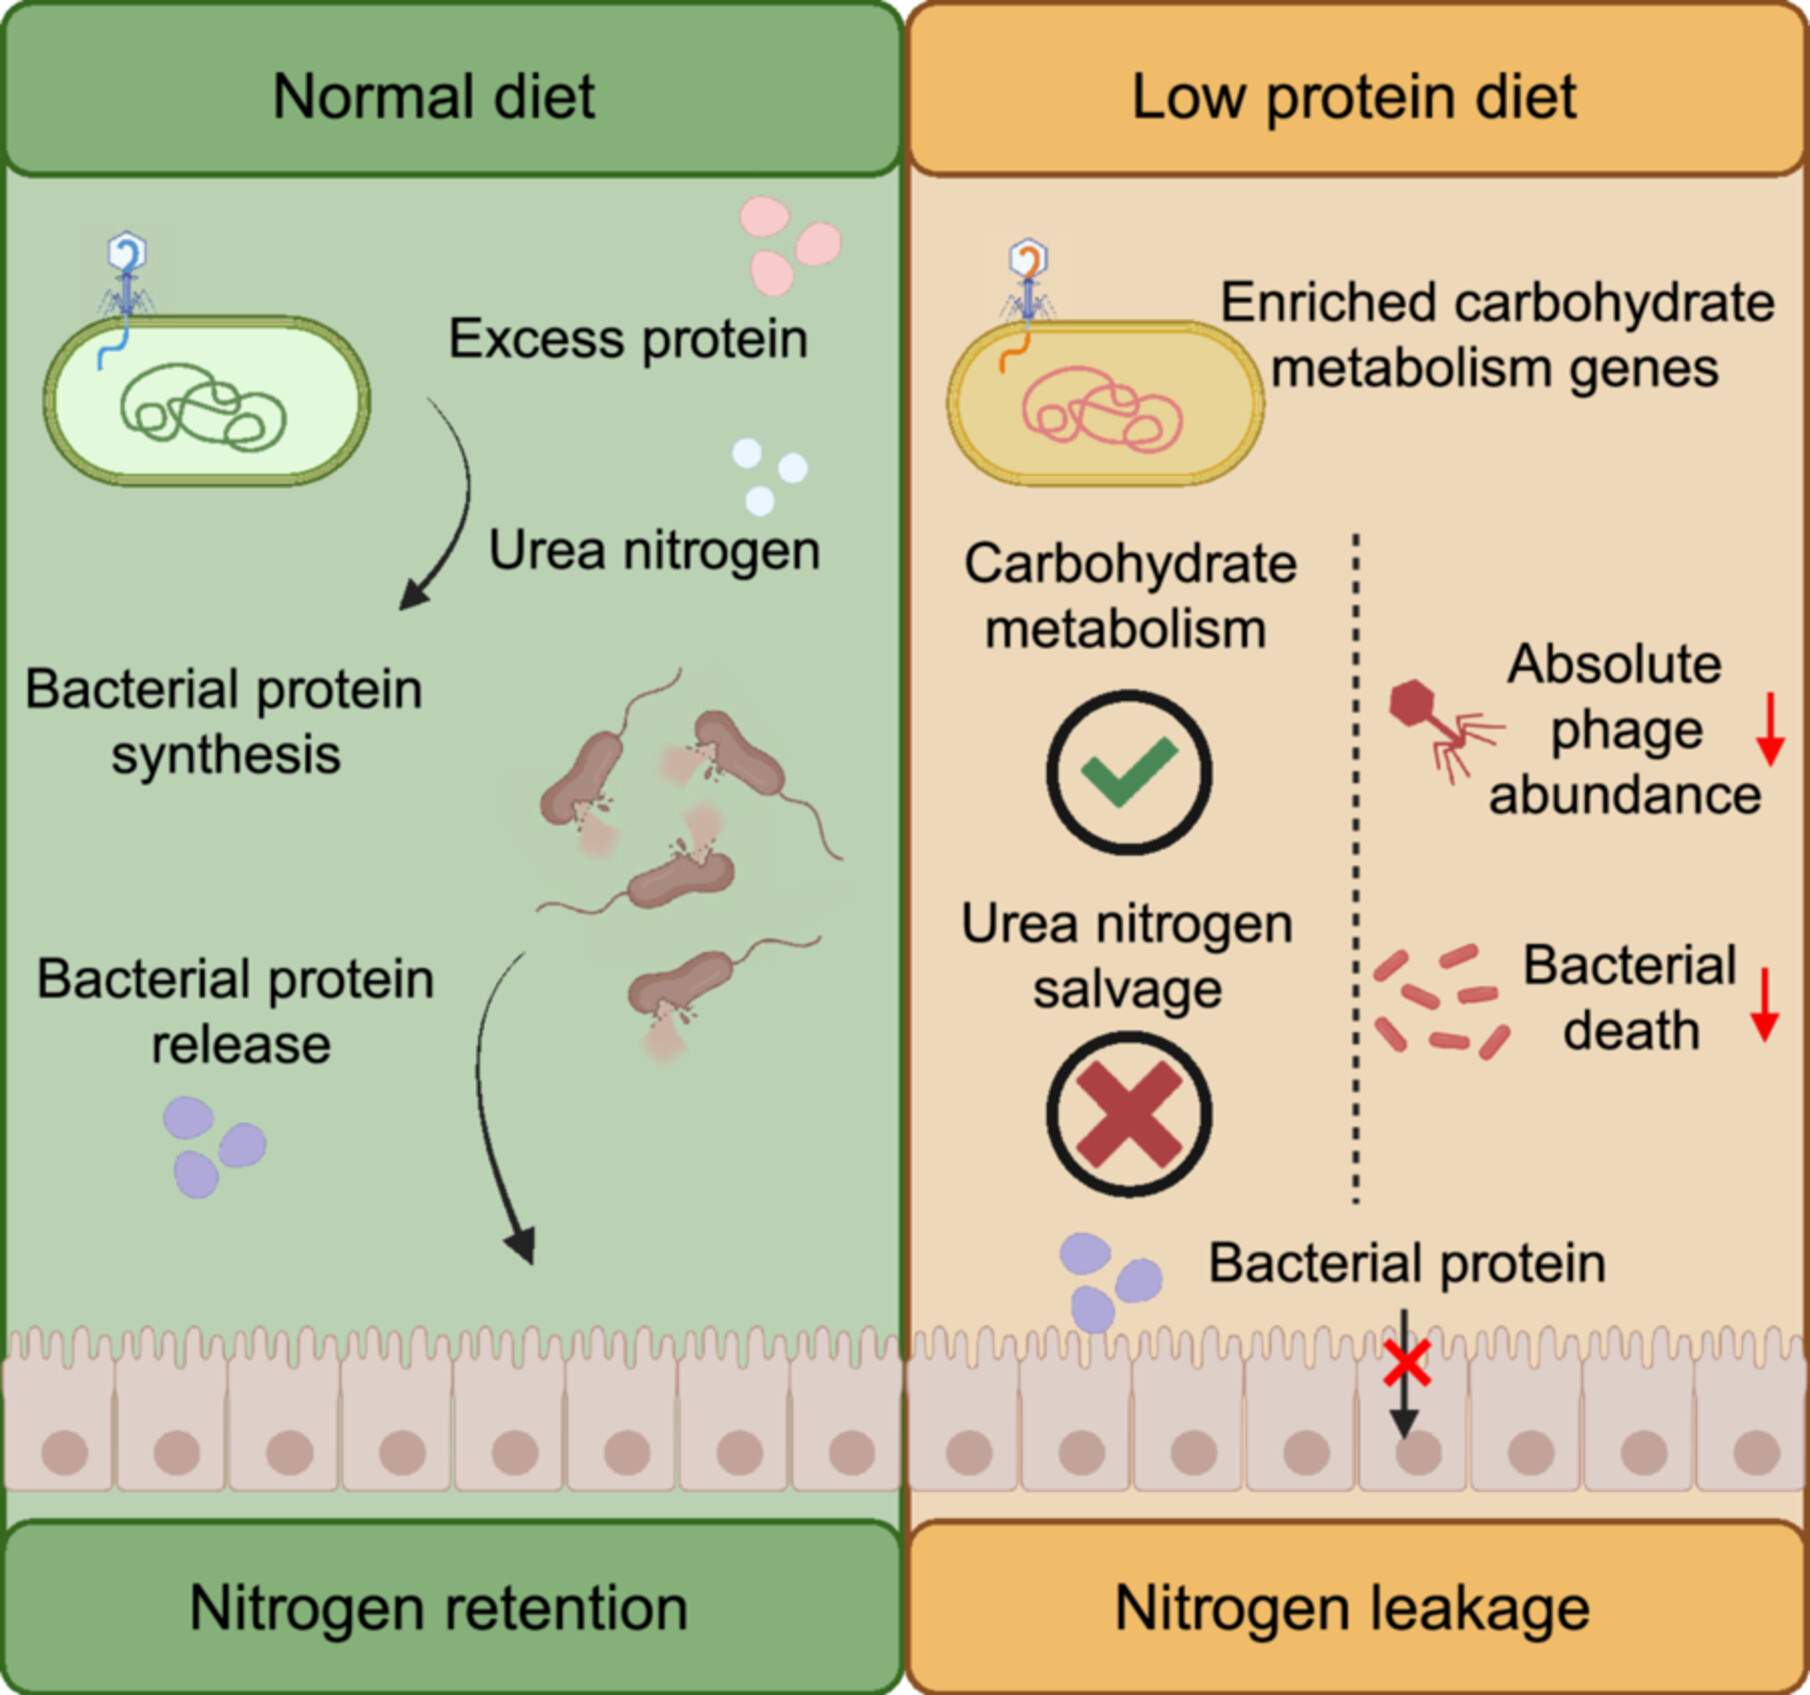
Under normal protein intake, lytic phages promote bacterial turnover, releasing microbial nitrogen that can be assimilated by the host and thereby revealing phage-mediated lysis as a potential pathway for host nutrient acquisition. Under protein restriction, however, phages shift toward a lysogenic strategy, reducing bacterial death and microbial protein release and ultimately limiting nitrogen availability to the host. Concurrently, protein restriction promotes the expansion of carbohydrate-utilizing bacteria in association with phages carrying carbohydrate metabolism-associated auxiliary metabolic genes, which further constrains ureolytic bacterial niches and thereby impairs intestinal urea nitrogen recycling.
Soil-derived, gut-dominant generalist bacteria shape the fitness of folivorous larvae
- 30 August 2026
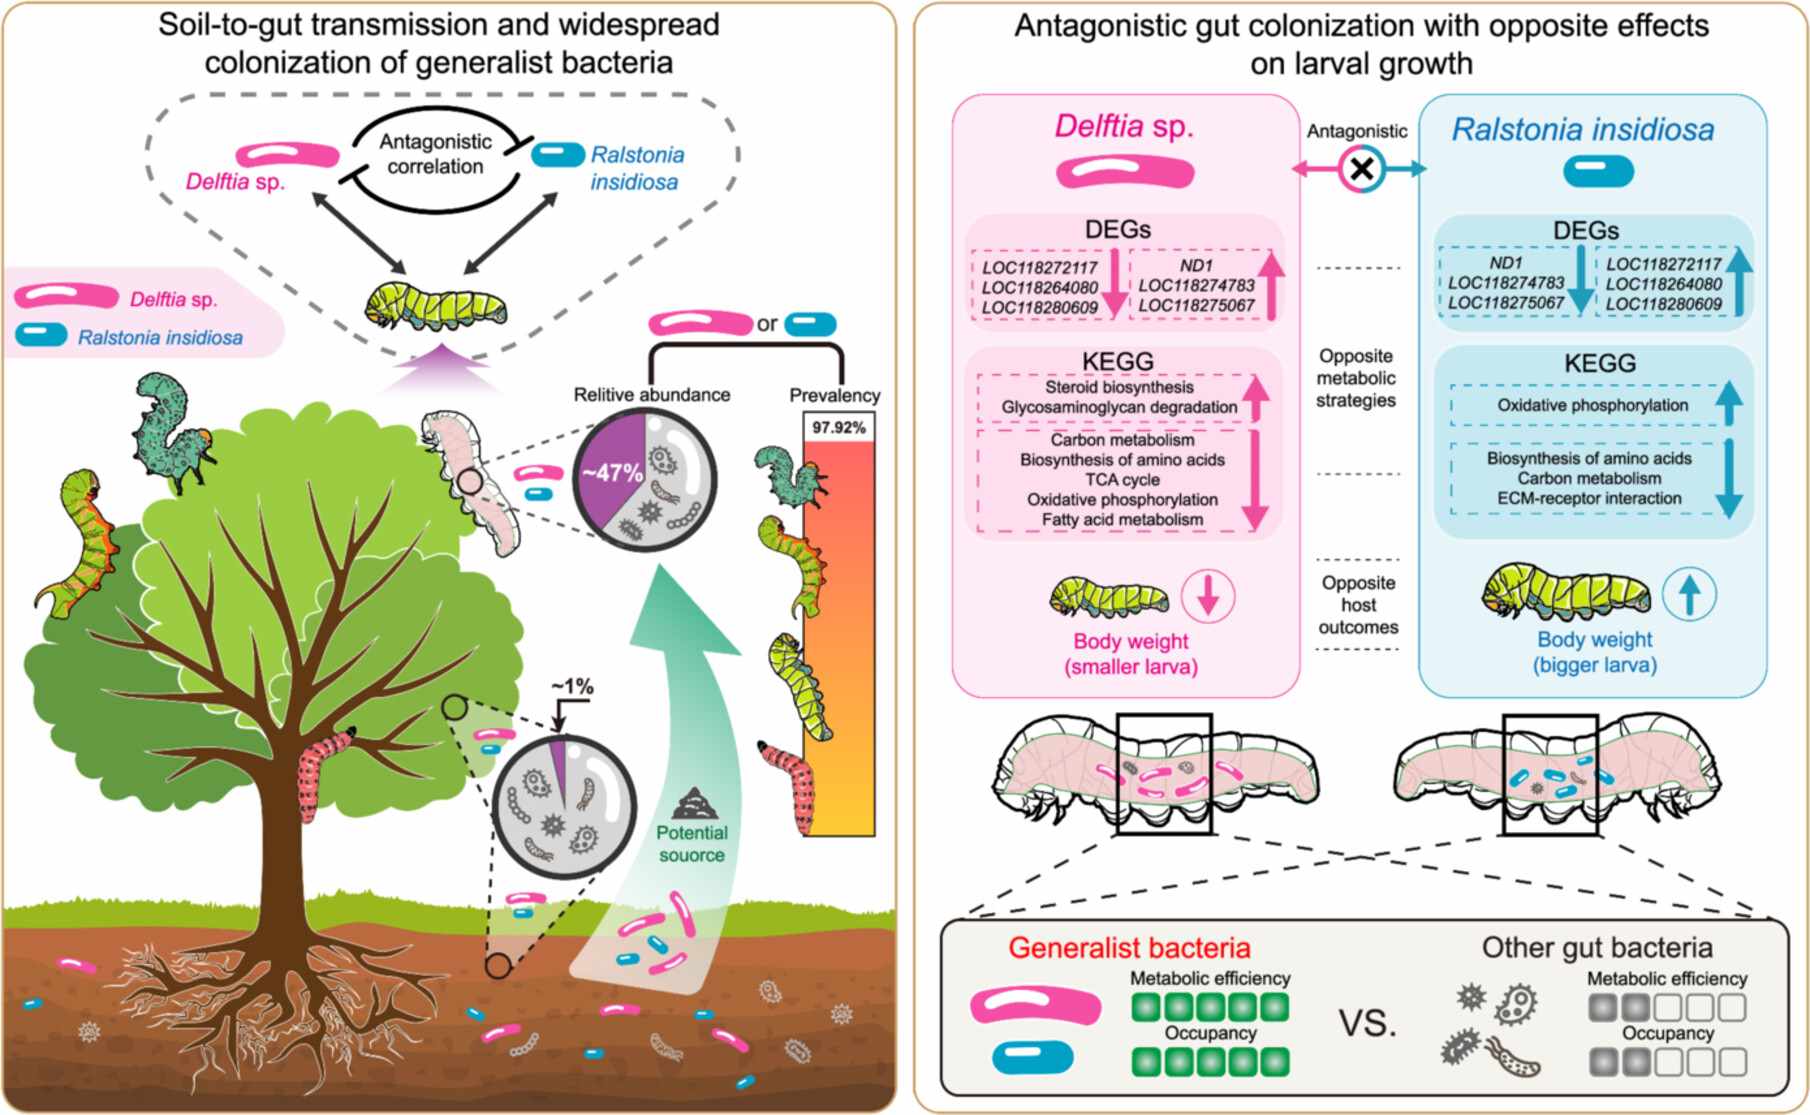
Whether Lepidoptera harbor a conserved core gut microbiome has long remained contentious. Through large-scale microbiome profiling of folivorous larvae, their host plants, and associated soils across three climatically distinct regions of China, we identify two soil-derived generalist bacteria, Ralstonia insidiosa and Delftia sp., that colonize 97.92% of larval species examined, attaining mean relative abundances exceeding 47%, with the soil microbial reservoir as their principal source. Strikingly, these two taxa exhibit strong mutual exclusion within the larval gut yet govern host development through diametrically opposed metabolic strategies: R. insidiosa promotes larval weight gain, whereas Delftia sp. suppresses growth. This functional bifurcation, in which two widespread generalists exert opposite phenotypic effects, represents a previously undescribed phenomenon in insect-microbe symbiosis. Our findings provide broad evidence that soil microbial reservoirs can shape aboveground herbivore fitness via horizontally acquired bacteria, offering mechanistic insights for microbiome-based ecological management.
Infection mechanisms of Rhizoctonia cerealis in wheat
- 05 October 2026
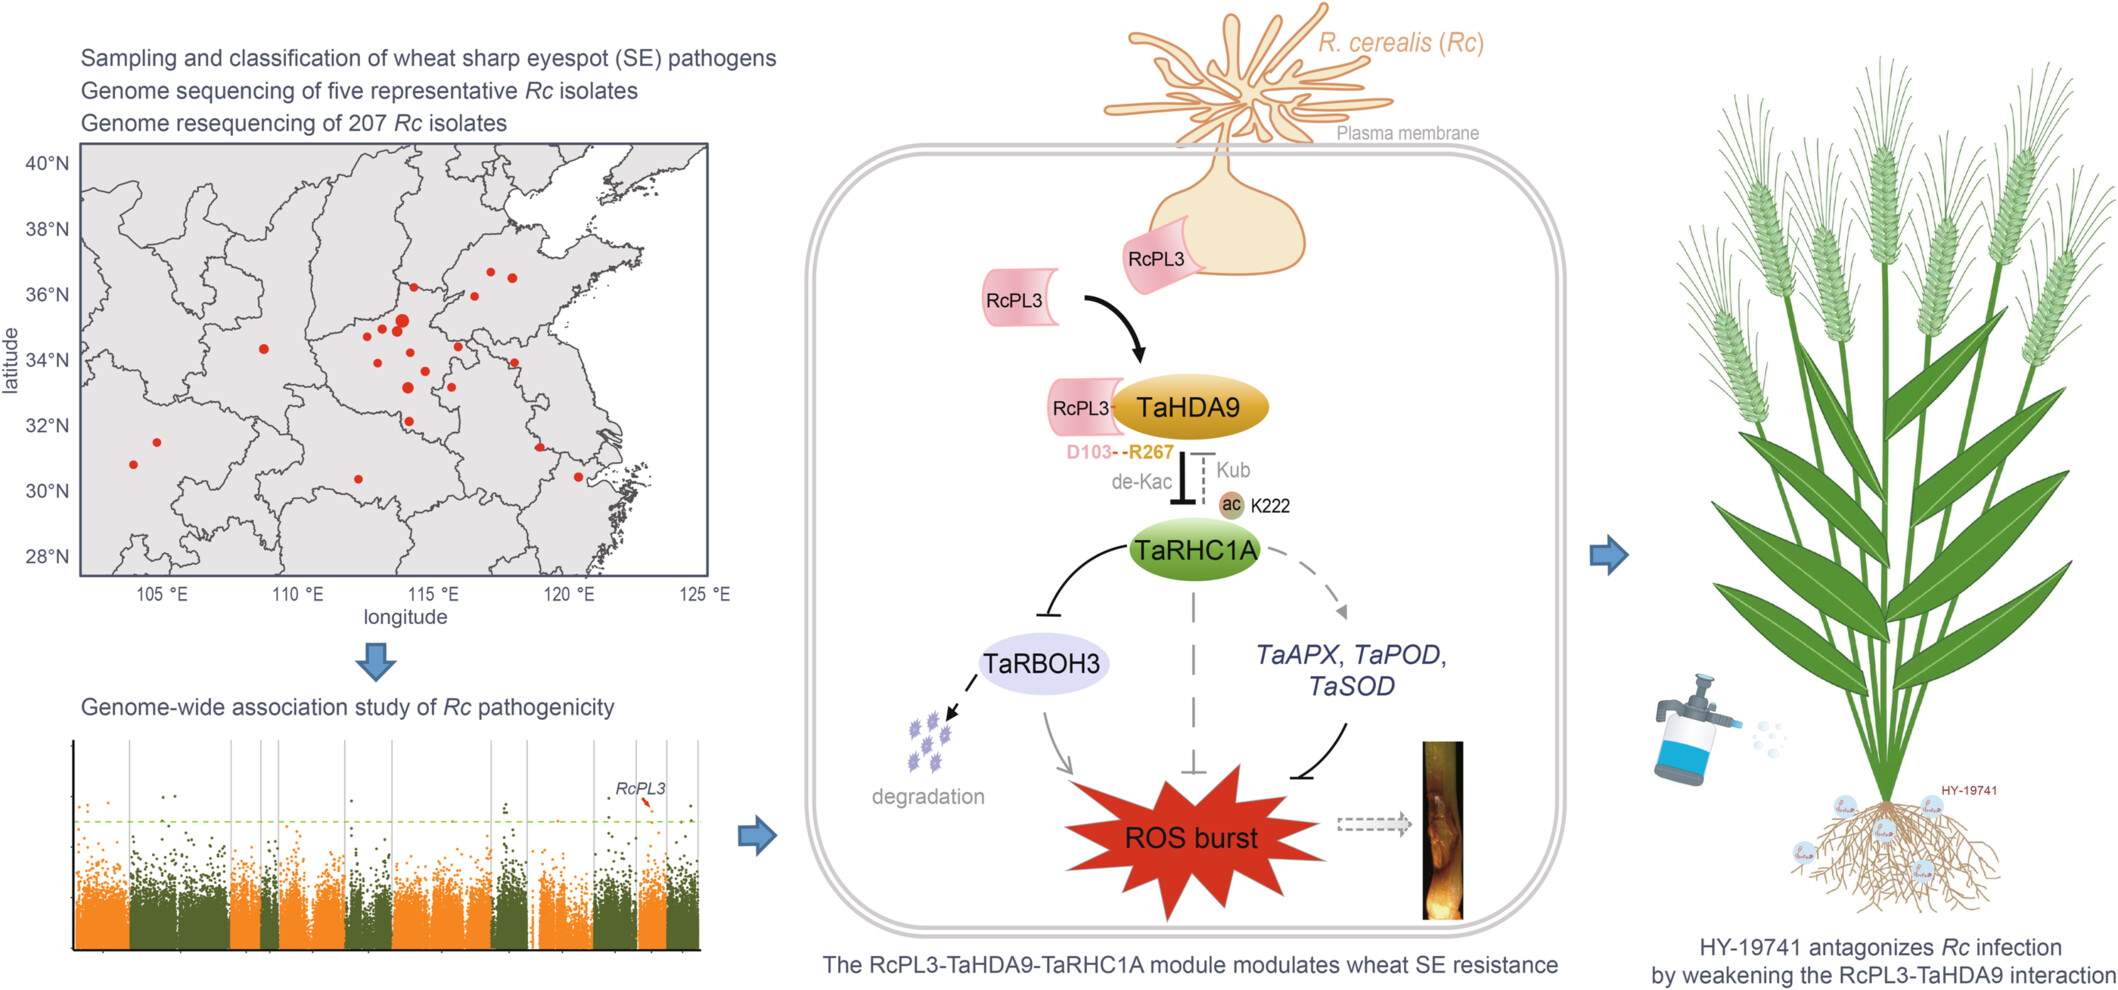
We discover that R. cerealis (Rc) is a primary pathogen causing sharp eyespot of wheat after 335 distinct isolates were isolated from 1683 SE-diseased wheat plants across China over 7 years using a single hyphal tip isolation method. We then generated high-quality reference genomes via de novo sequencing of five representative Rc isolates and constructed a genomic variation atlas by resequencing 207 Rc isolates. We further identify RcPL3 as a key effector by integrating genome, transcriptome, and genome-wide association study analyses, and find that RcPL3 recruits TaHDA9 to suppress TaRHC1A by deacetylation and thus inhibits degradation of TaRBOH3 by TaRHC1A, finally resulting in ROS burst that contributed to SE susceptibility in wheat plants. Furthermore, we identified HY-19741 as an antagonist to effectively impede Rc infection by weakening the interaction of RcPL3 and TaHDA9. This work reveals a novel fungal pathogenic cascade and provides a candidate agent for wheat disease control.
Deciphering transcriptome complexity via long-read sequencing
- 04 October 2026
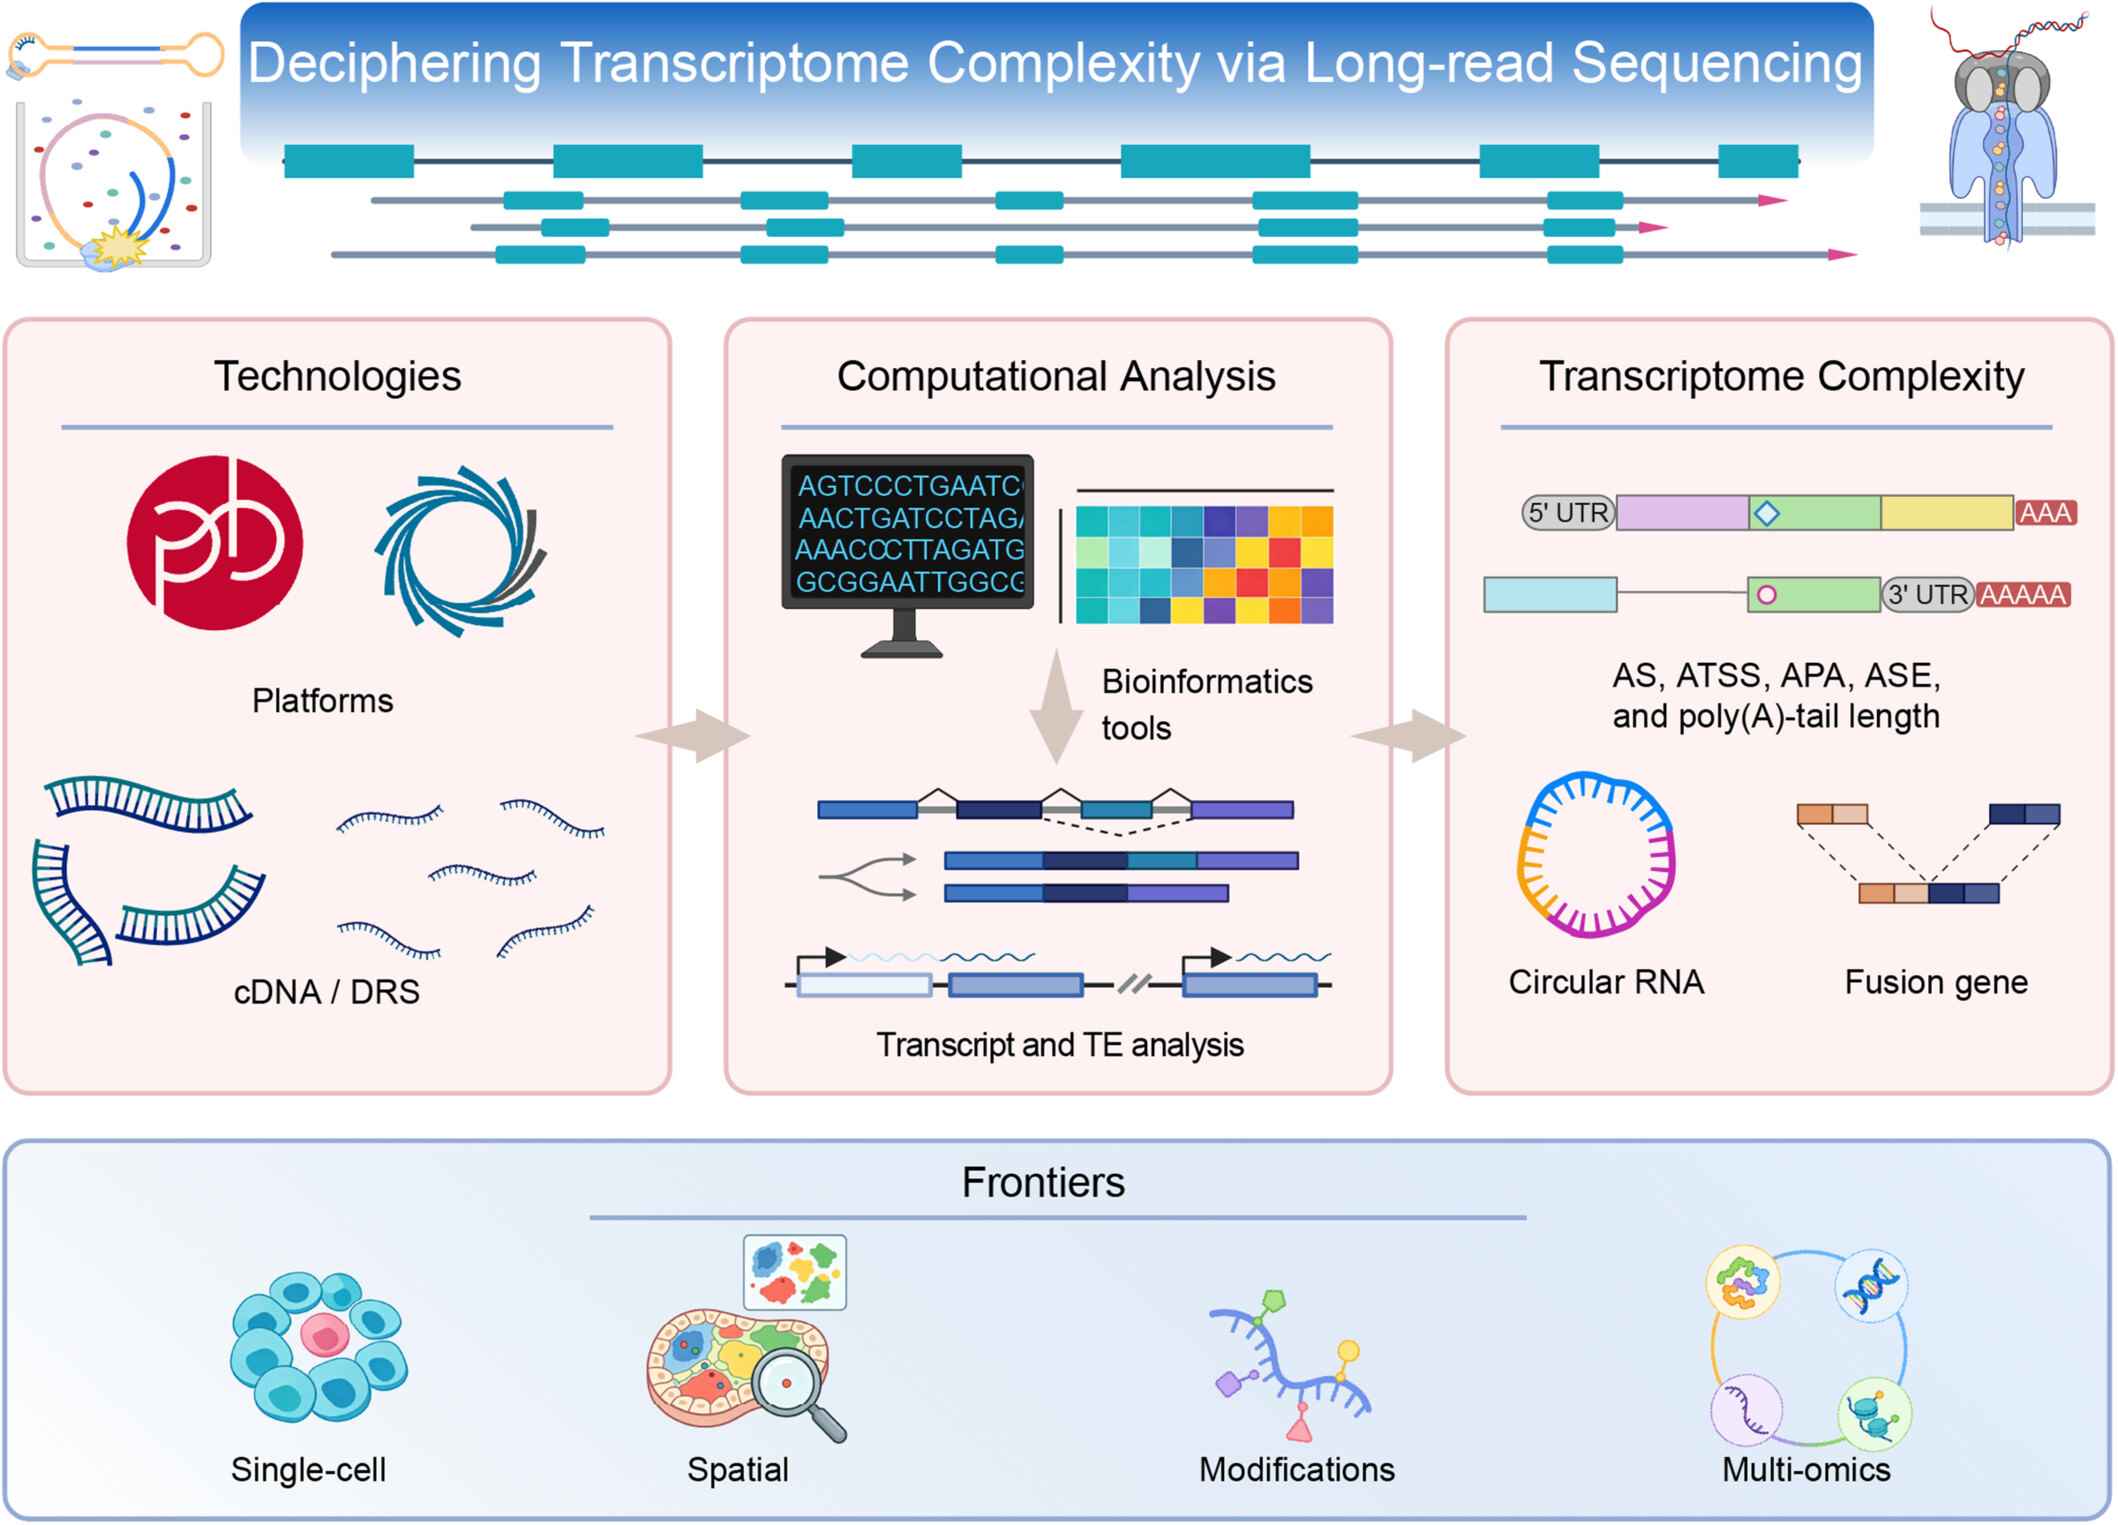
Long-read sequencing is transforming transcriptomics from gene-level quantification to isoform-resolved interpretation by directly resolving full-length transcript structures, transcription in repetitive regions, and linked molecular features. This review provides an end-to-end roadmap covering sequencing platforms, library preparation strategies, computational workflows, the analysis of transcriptome complexity, and downstream analyses. It further highlights current challenges and emerging frontiers, including single-cell and spatial transcriptomics, direct RNA modification detection, and integrated multi-omics approaches.
Monocyte inflammation and adaptive immune dysfunction in severe Klebsiella pneumoniae pneumonia
- 21 September 2026
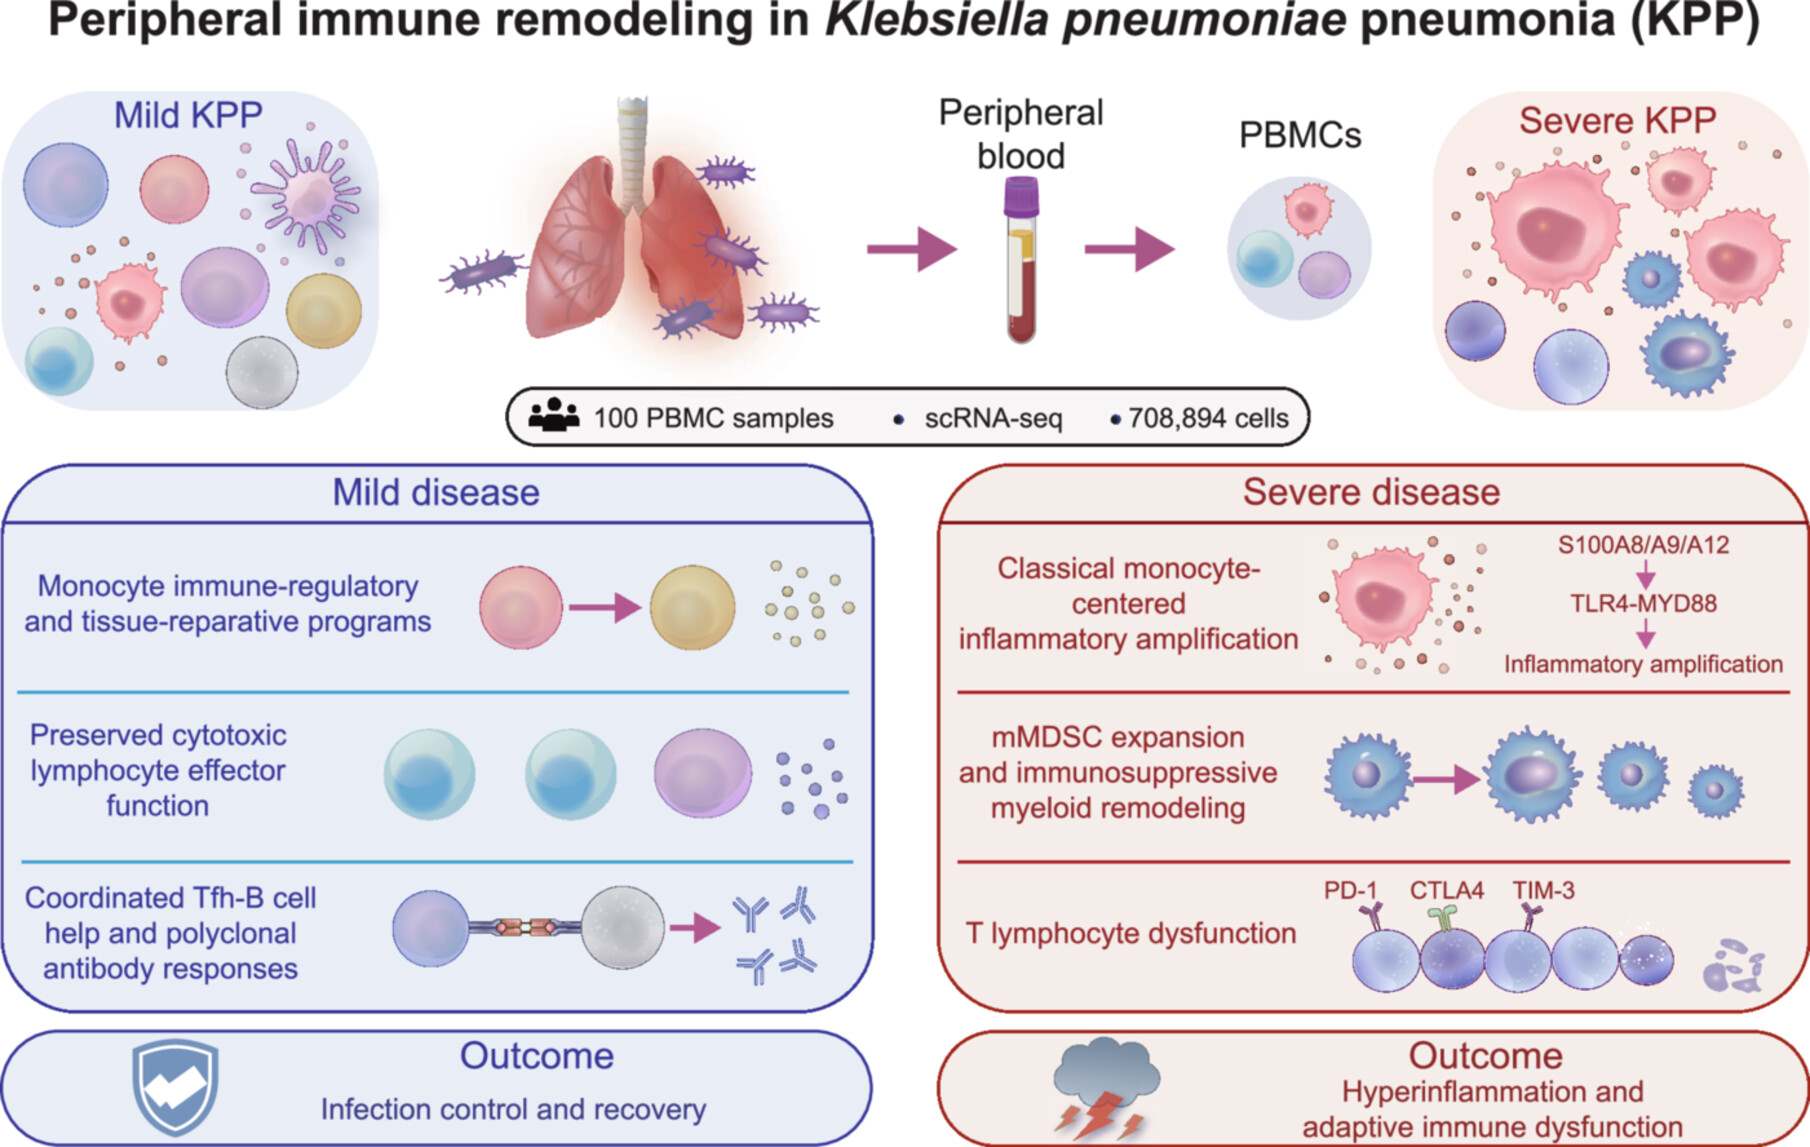
Severe Klebsiella pneumoniae pneumonia combines intense inflammation with failure to control infection, yet the cellular basis is unclear. Using a severity-focused single-cell atlas of 708,894 peripheral blood mononuclear cells from 100 individuals, complemented by plasma protein measurements and flow cytometry, we show that disease severity redirects rather than simply amplifies systemic immunity. Mild disease preserves coordinated host defense: immune-regulatory and tissue-reparative monocytes, intact cytotoxic lymphocytes, and T follicular helper–B cell coordination. Severe disease is characterized by S100A8/A9/A12-high inflammatory monocytes with enhanced TLR4–MYD88 signaling, expansion of monocytic myeloid-derived suppressor cell-like states, and impaired dendritic cell antigen presentation. These myeloid changes accompany dysfunctional CD4+ T cells, cytotoxicity- and exhaustion-associated CD8/NK cells, and weakened expression of B cell receptor signaling-related genes despite plasma cell accumulation. Severe disease is characterized by a systemic pattern of impaired immune coordination, highlighting monocyte-derived S100 alarmins as candidates for future mechanistic and therapeutic investigation.
Gut Enterococcus faecium derived leucic acid mediates the antiobesity effect of dietary fiber
- 21 September 2026
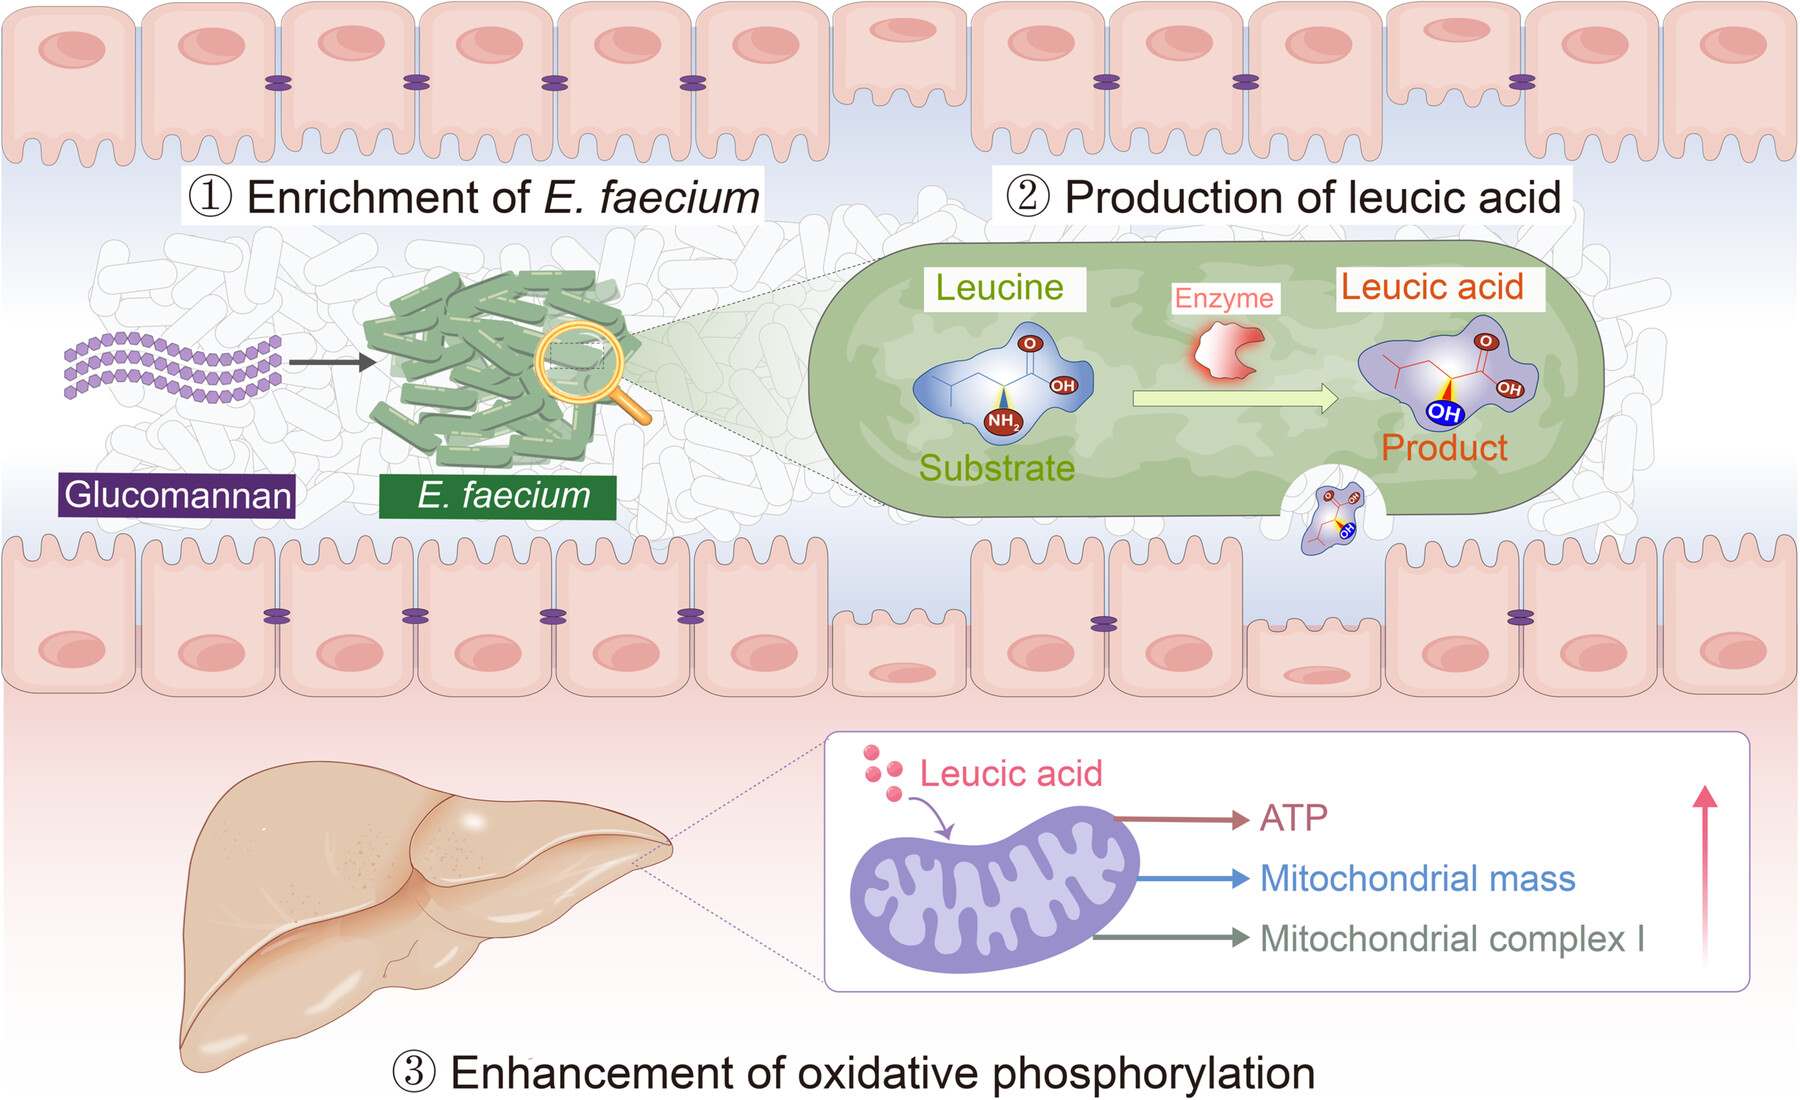
This study identified Enterococcus faecium as a gut bacterium with anti-obesity activity, which alleviated adipocyte hypertrophy, dyslipidemia, and hepatic steatosis by converting leucine into leucic acid (LEA) to regulate mitochondrial oxidative phosphorylation. In addition, dietary glucomannan effectively enriched this bacterium, providing a theoretical basis for developing precision dietary fiber-based anti-obesity strategies targeting the gut microbiota.
Decoding sequence recognition code of nucleic acid-binding proteins of human-infecting DNA viruses
- 08 September 2026
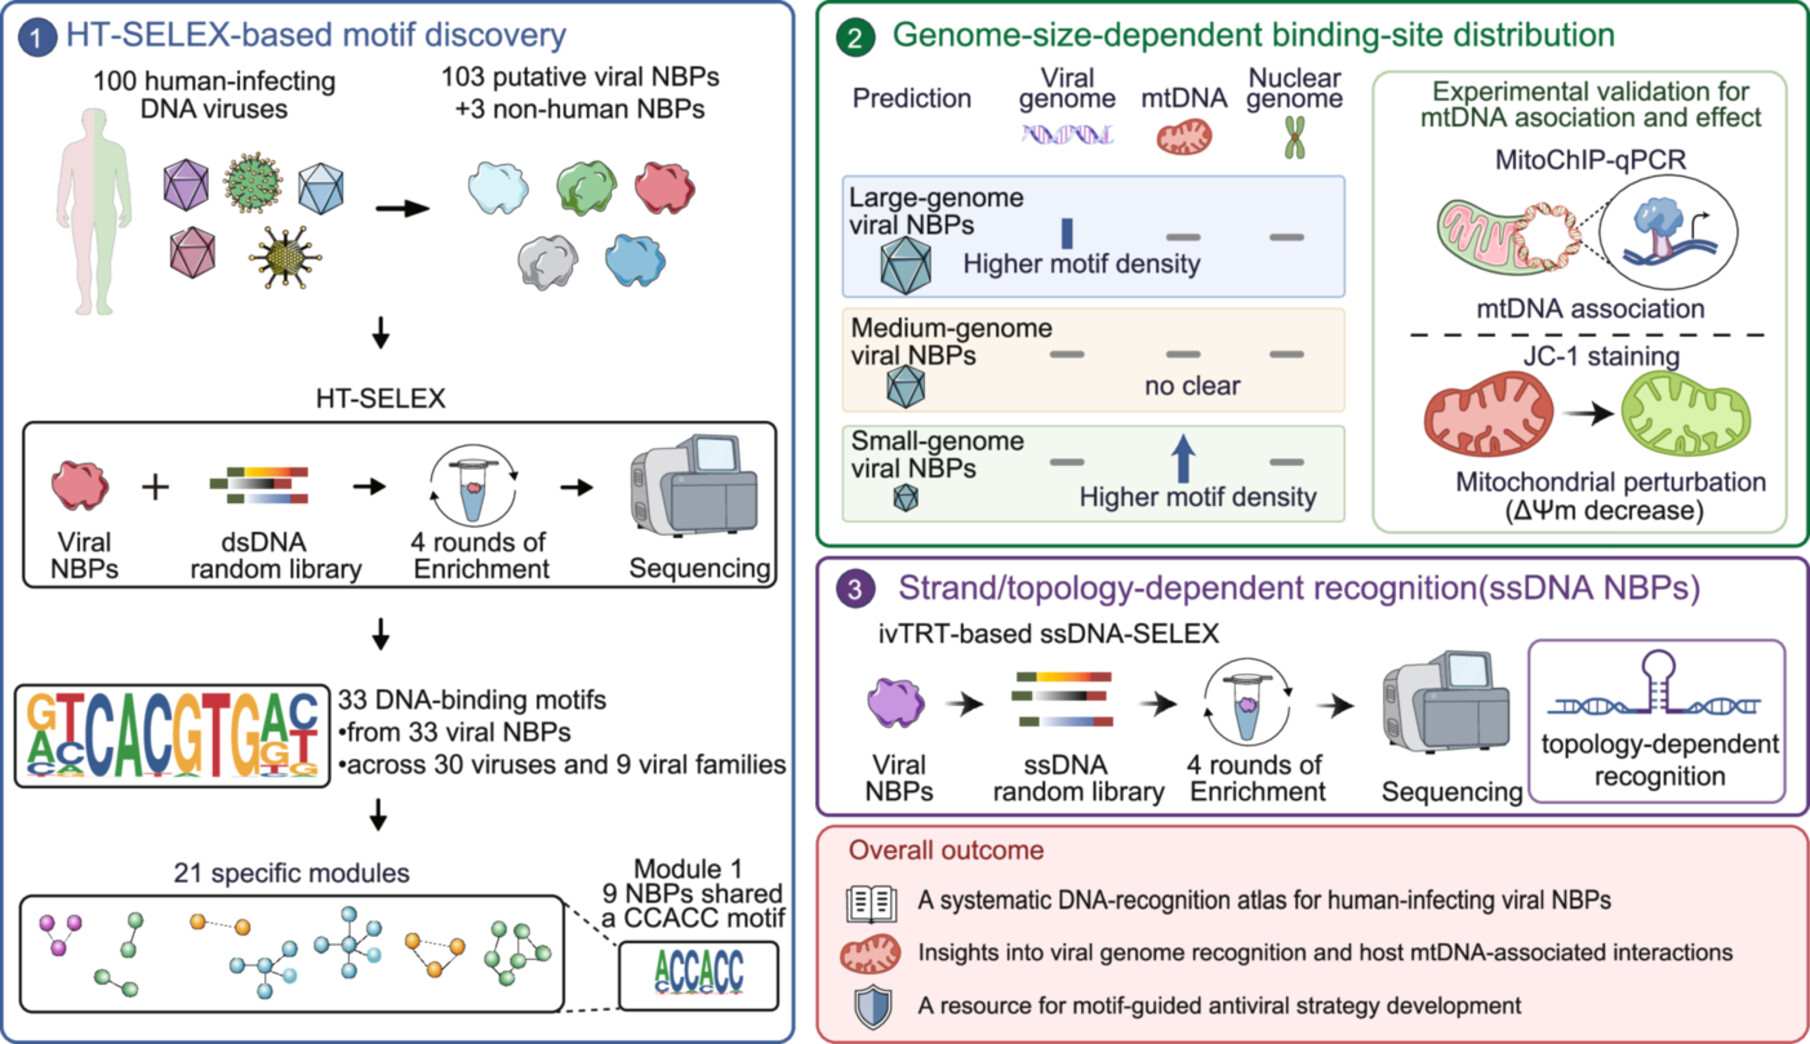
Human-infecting DNA viruses remain major health threats, yet the DNA-recognition mechanisms of their nucleic acid-binding proteins (NBPs) are poorly understood. Here, we systematically profiled 103 viral NBPs from human-infecting DNA viruses, with three NBPs from non-human-infecting DNA viruses as controls, using high-throughput screening. This analysis identified diverse DNA-binding motifs and specificity modules, including convergent recognition of a conserved CCACC motif across phylogenetically distant viruses. Notably, viral NBP binding-site distributions varied with genome size, and several NBPs from small-genome viruses showed enrichment on mitochondrial DNA. Functional assays further supported their mitochondrial association and effects on mitochondrial membrane potential. By integrating an ivTRT-based ssDNA-SELEX workflow, we further found that ssDNA viral NBPs recognize dimer-like and inverted-repeat sequences with potential to form stem-loop structures. Collectively, this study constructs a comprehensive viral NBP DNA-recognition atlas, offering a fundamental resource for elucidating viral genome recognition mechanisms, virus–mitochondria interactions, and developing future antiviral strategies.
Human biomarker navigator
- 20 August 2026

The Human Biomarker Navigator integrates the disease continuum, biomarker dynamics, cross-organ biomarker networks, biomarker classification, and technology-driven paradigms. It maps how biomarkers link multi-system physiology and pathology across the nervous, respiratory, endocrine, circulatory, immune, digestive, urinary, reproductive, and musculoskeletal systems. Within the cross-organ network, representative biomarkers span inflammation and tissue remodeling (SAA, PTX3, MMPs), pulmonary function (FEV1/FVC), angiogenesis and vascular injury (VEGF, PlGF), oncology and minimal residual disease (cfDNA, CTC, MRD, FLT3-ITD, KRAS, PD-L1, CEA, CA125, CA19-9, GPC1/GPC3), cardiac injury and heart failure (hs-cTn, CK-MB, NT-proBNP, Gal-3), hepatic metabolism and injury (ALT, AST, GGT, ALP, DCP, bile acids), renal function and damage (eGFR, UACR, cystatin C, KIM-1, NGAL), neuroimmunity (Aβ42/40, MOG-IgG, LGI1 antibodies), and bone turnover (PINP, OPG, OC, β-CTX, DKK-1). By enabling integrative detection, it supports early diagnosis, stratification, and treatment monitoring.
Multi-omics–driven precision medicine
- 25 August 2026
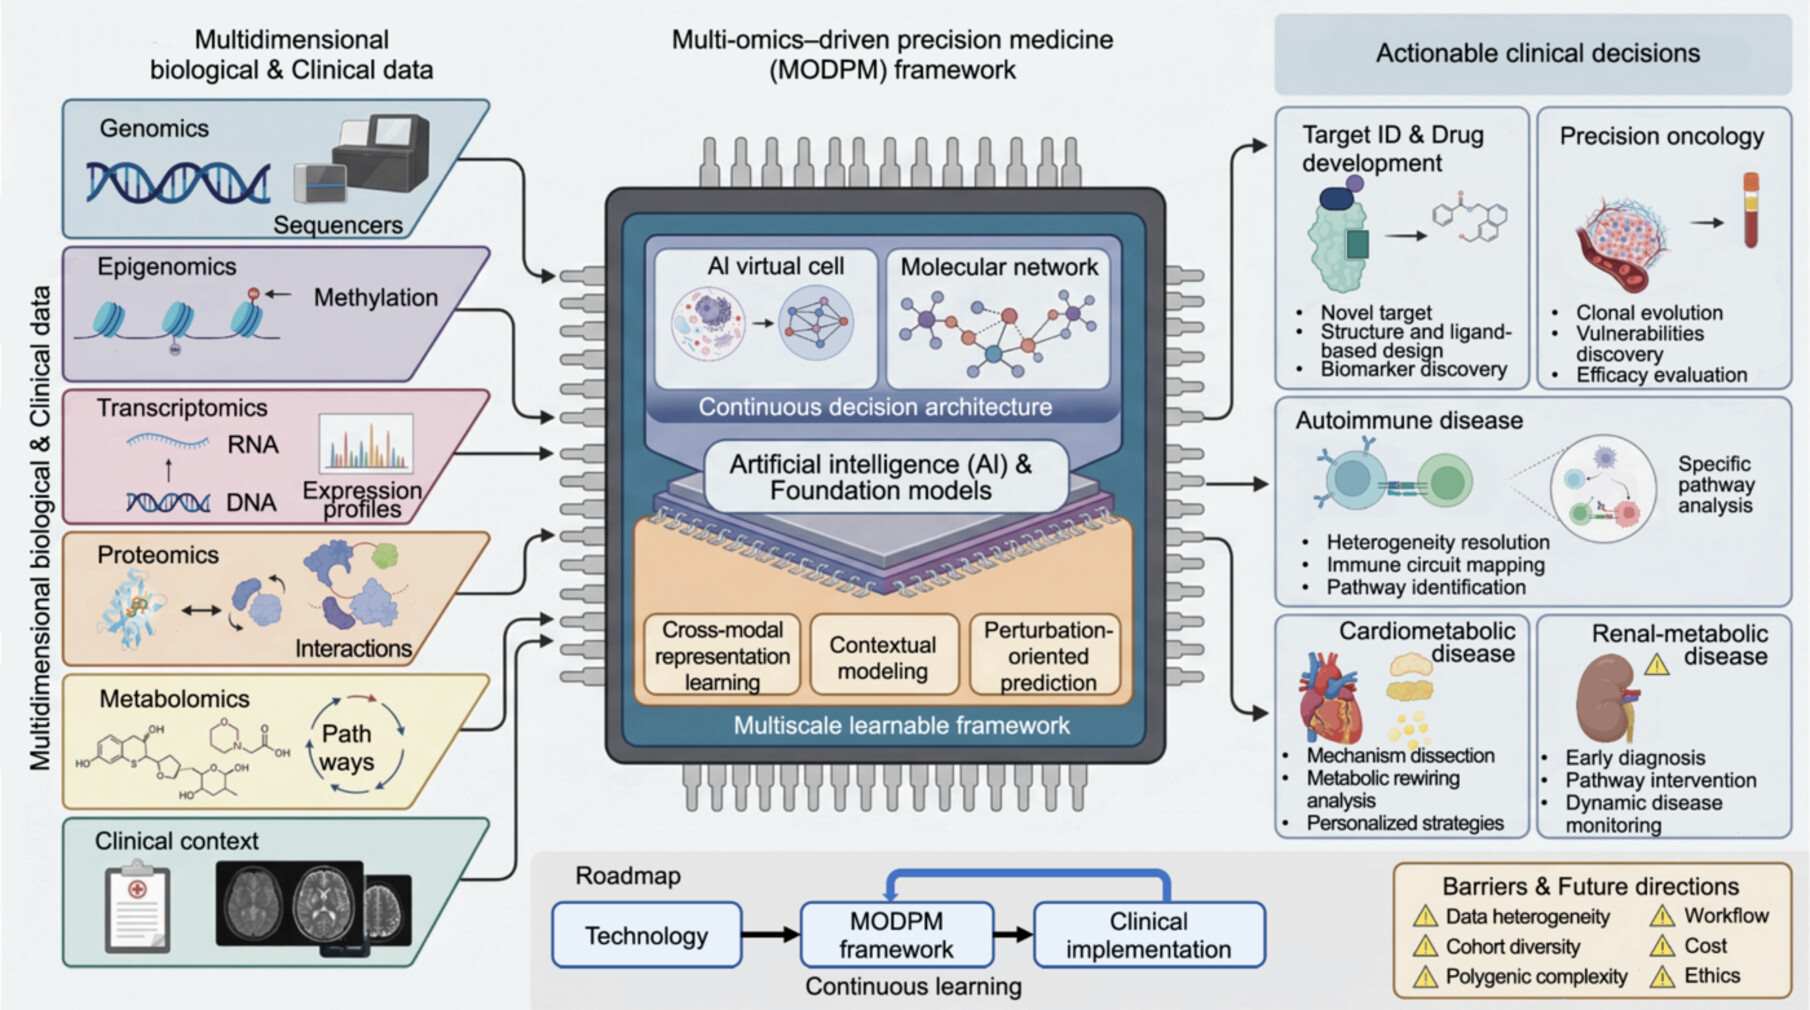
Multi-omics-driven precision medicine (MODPM) provides a multiscale, continuously learnable framework that integrates genomics, epigenomics, transcriptomics, proteomics, metabolomics, microbiome profiles, and clinical data. Powered by artificial intelligence and foundation models, MODPM enables cross-modal representation learning, contextual modeling, molecular network inference, and perturbation-oriented prediction. This framework transforms multidimensional biological information into actionable clinical decisions by advancing target identification, drug development, biomarker discovery, disease stratification, precision oncology, and individualized management across disease areas such as autoimmune, cardiometabolic, and renal–metabolic diseases. Continuous feedback from clinical implementation further refines models, supports dynamic monitoring, and addresses barriers related to heterogeneity, cohort diversity, workflow, cost, and ethics.
Beyond omics: From descriptive profiling to causal and scalable design of fermentation microbiomes
- 30 July 2026
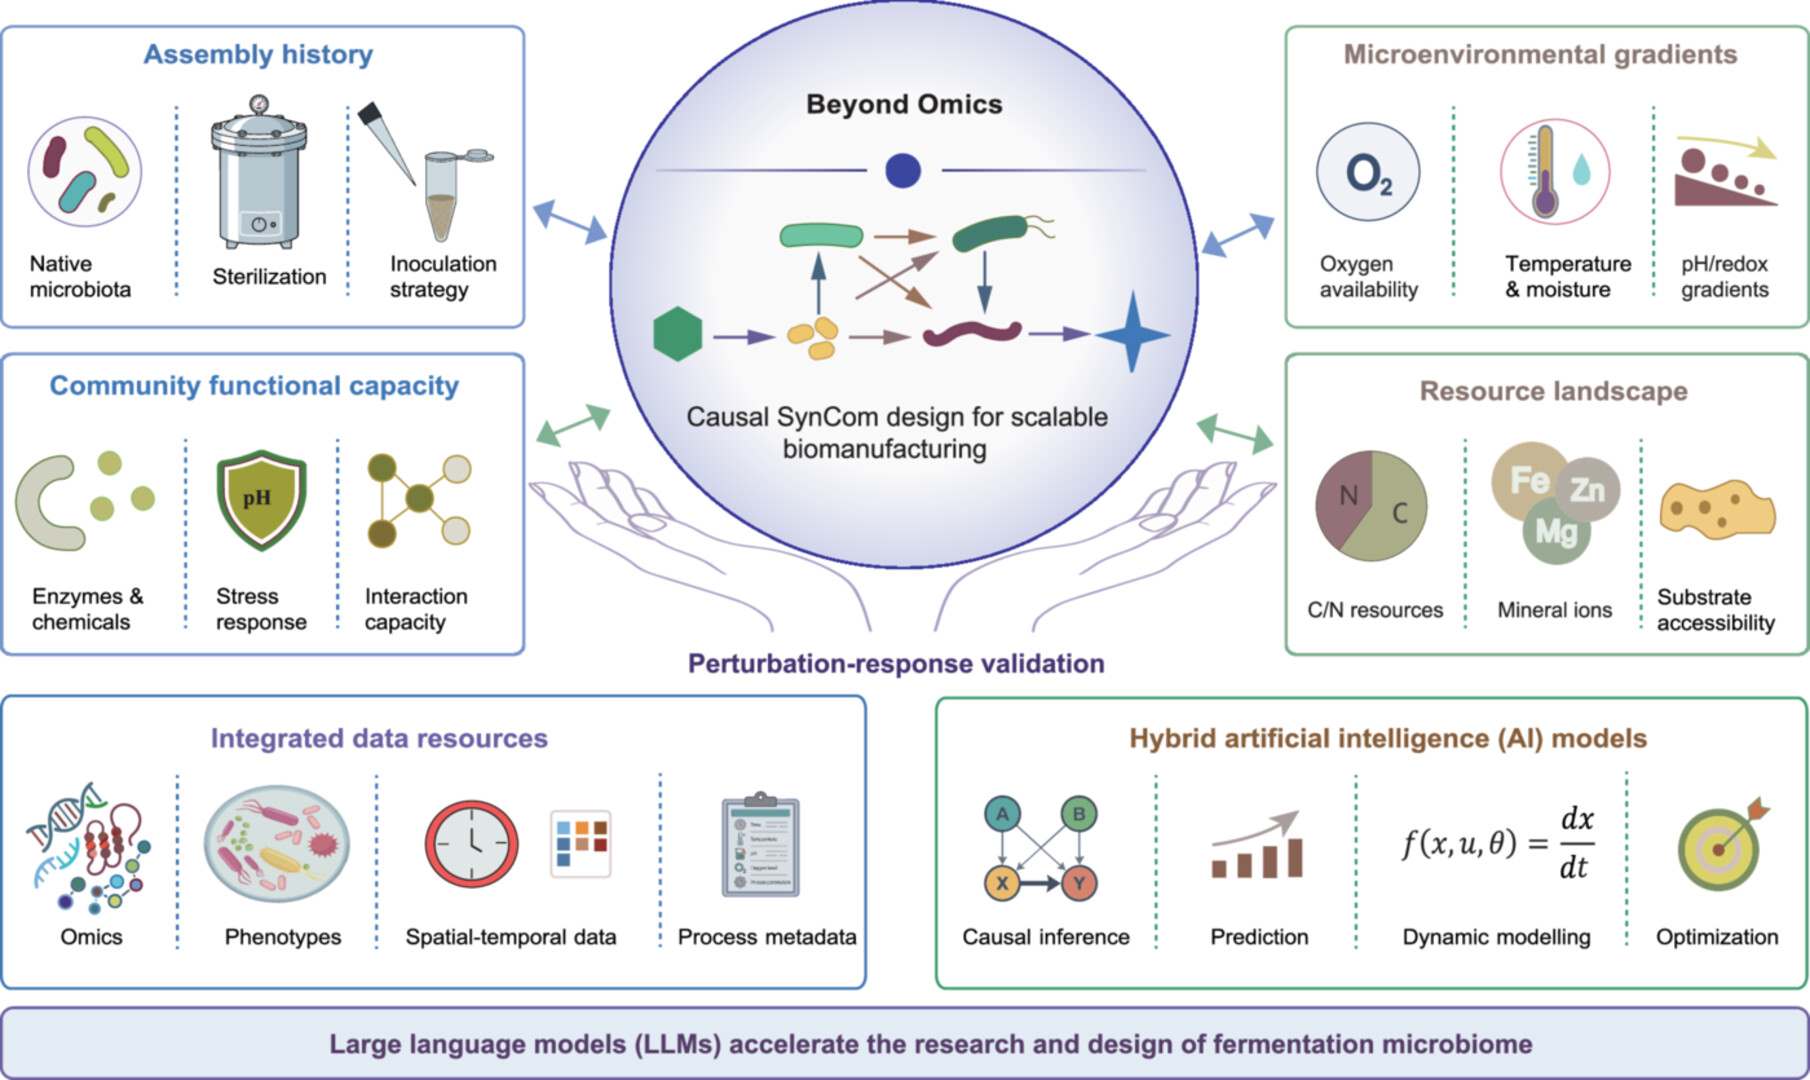
Fermentation microbiomes play essential roles in food production, feed preservation, waste valorization, and diverse sustainable industrial processes. Although multi-omics and systems biology have substantially advanced our understanding of their assembly, interactions, and functional dynamics, industrial translation remains constrained by fragmented datasets, limited causal validation, and transport constraints during scale-up. Large language models act as upper-level knowledge and workflow orchestrators, accelerating data integration, hybrid AI-mechanistic modeling, hypothesis generation, and perturbation-guided learning. Collectively, these advances enable fermentation microbiome research to move beyond descriptive omics toward mechanism-guided synthetic microbial community design, causal validation, and scalable biomanufacturing.
fastp 1.0: An ultra-fast all-round tool for FASTQ data quality control and preprocessing
- 09 September 2025
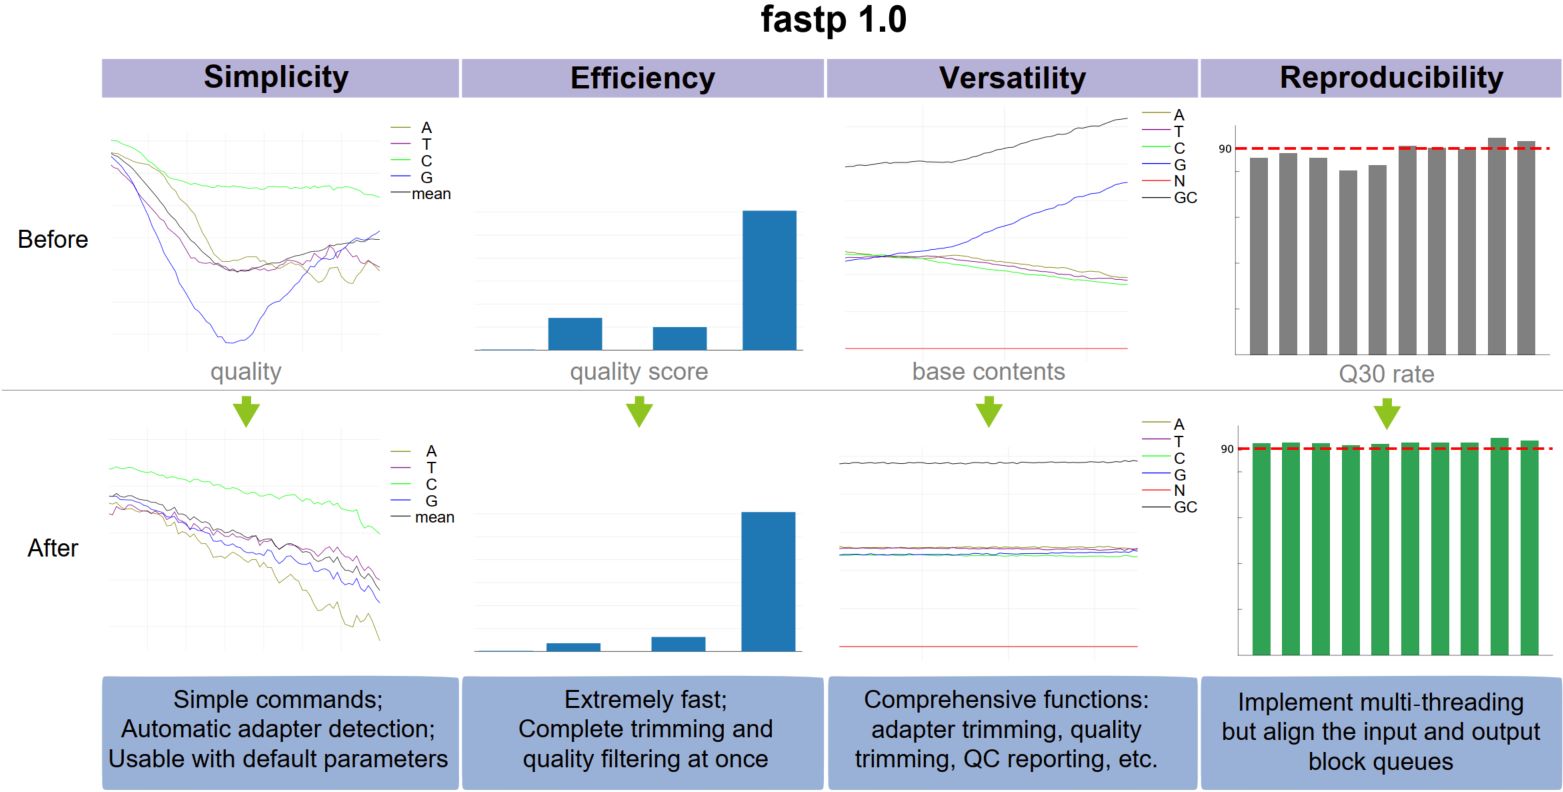
The figure shows side-by-side quality comparison of the data before and after preprocessing using fastp 1.0. As can be seen from this figure, the data quality has been significantly improved after fastp processing. The Q20 ratio and Q30 ratio increased, and the GC base ratio returned to balance.
EasyMetagenome: A user-friendly and flexible pipeline for shotgun metagenomic analysis in microbiome research
- 14 February 2025
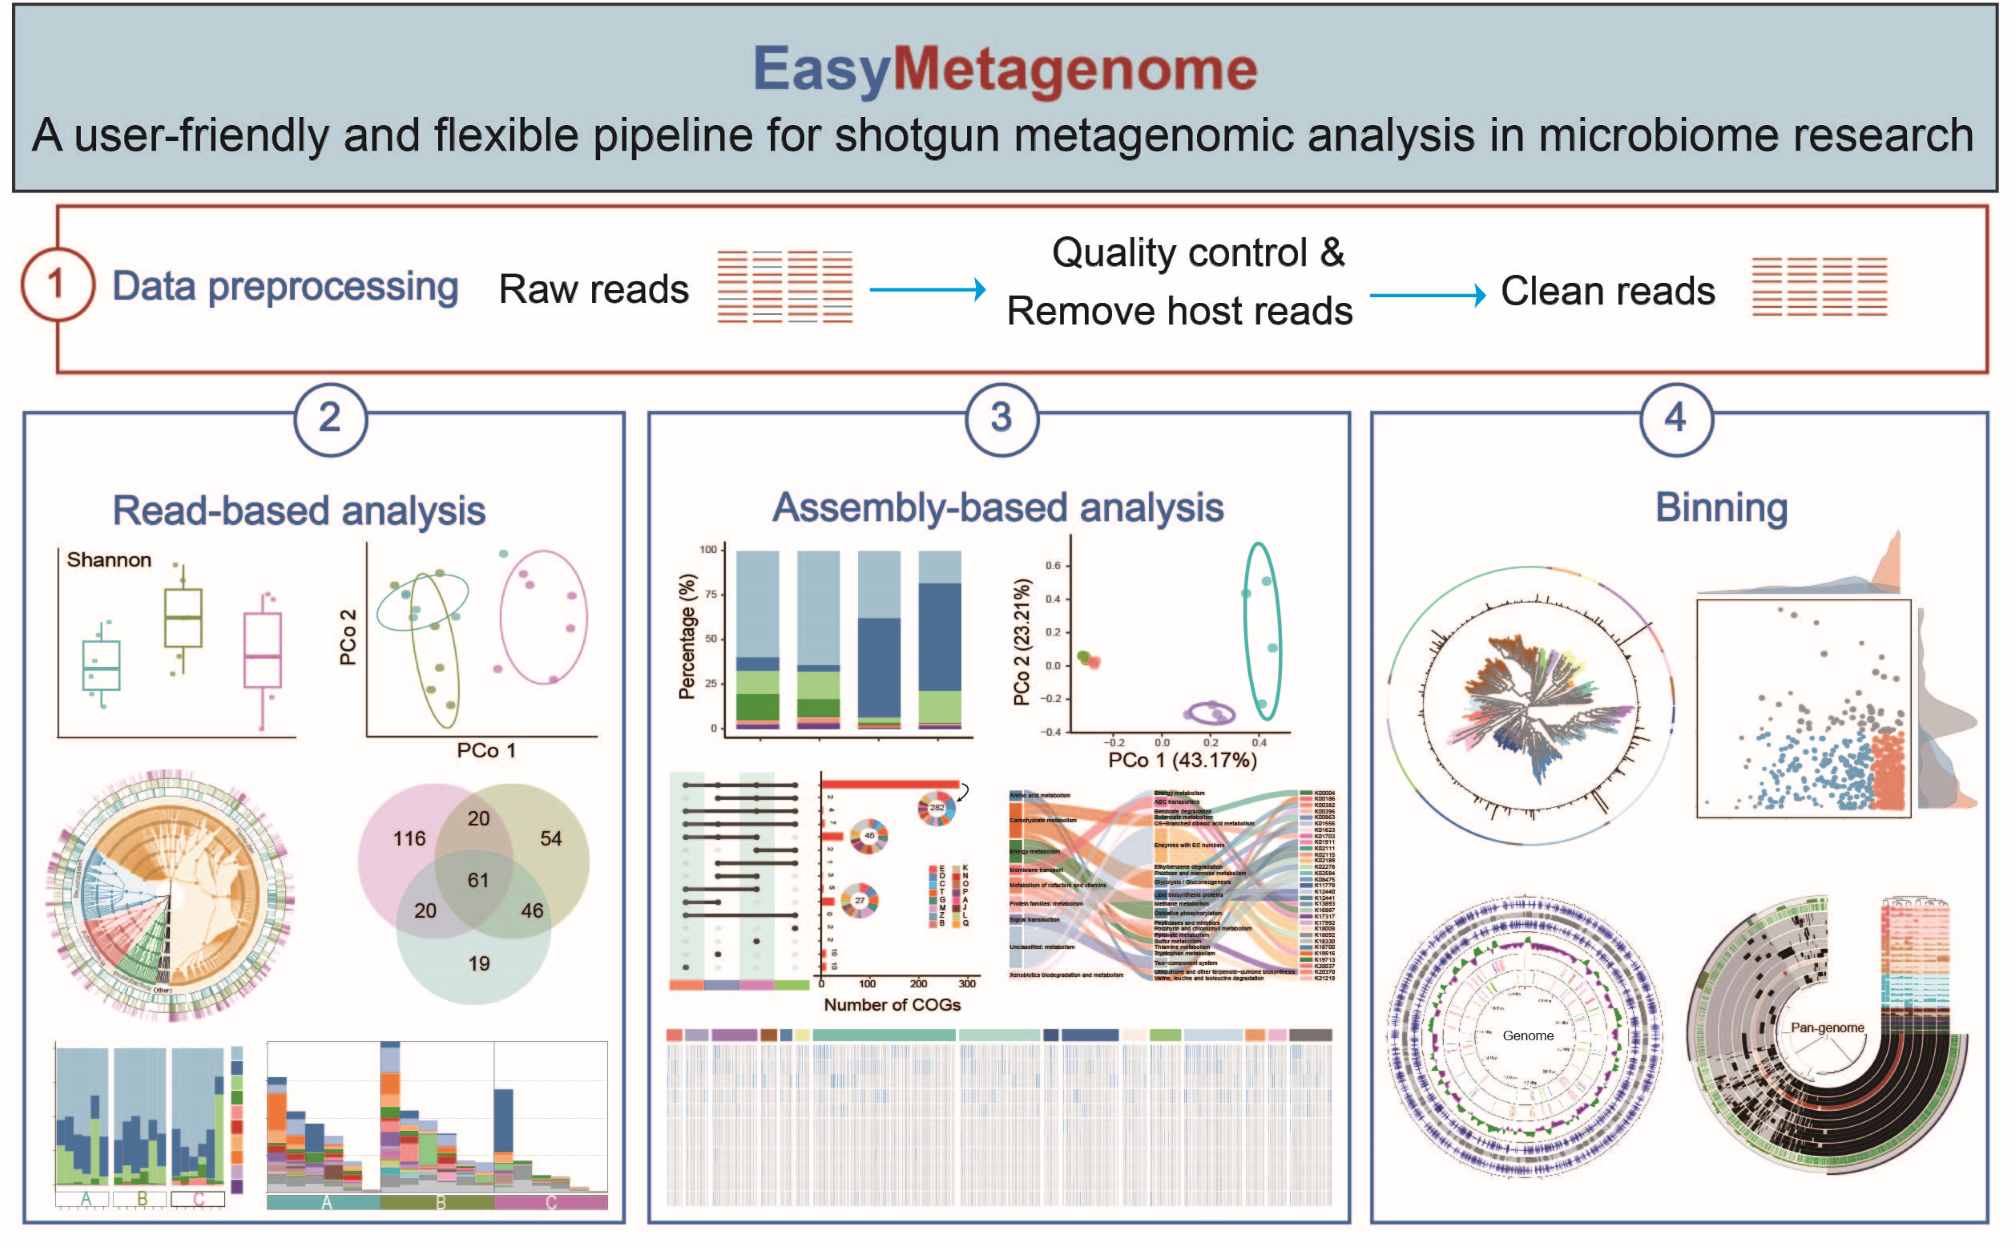
EasyMetagenome is a user-friendly shotgun metagenomics pipeline designed for comprehensive microbiome analysis, supporting quality control, host removal, read-based, assembly-based, binning, genome and pan-genome analysis. It offers customizable settings, data visualizations, and parameter explanations. The pipeline is freely available at https://github.com/YongxinLiu/EasyMetagenome.
Gut–X axis
- 26 February 2025
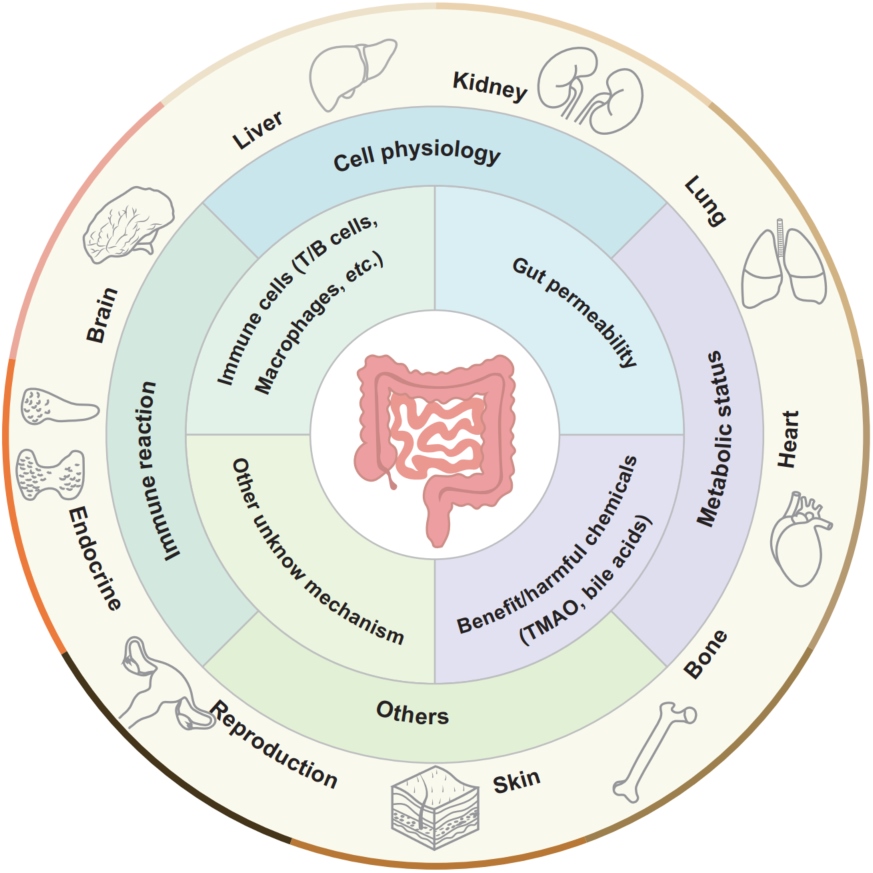
The concept of “gut–X axis”: the intestine and intestinal microbiota are proven to be able to modulate the pathophysiologic progressions of the extraintestinal organs' diseases. The bioactive chemicals and/or intestinal immune cells can translocate into the circulatory system and other organs and influence the immune reactions, metabolic status, cells physiology, and so forth of extraintestinal organs, finally regulating these organs' homeostasis. Meanwhile, other organs may reversely impact the intestine, namely such regulatory axis is bidirectional.
Ultrafast one-pass FASTQ data preprocessing, quality control, and deduplication using fastp
- 08 May 2023
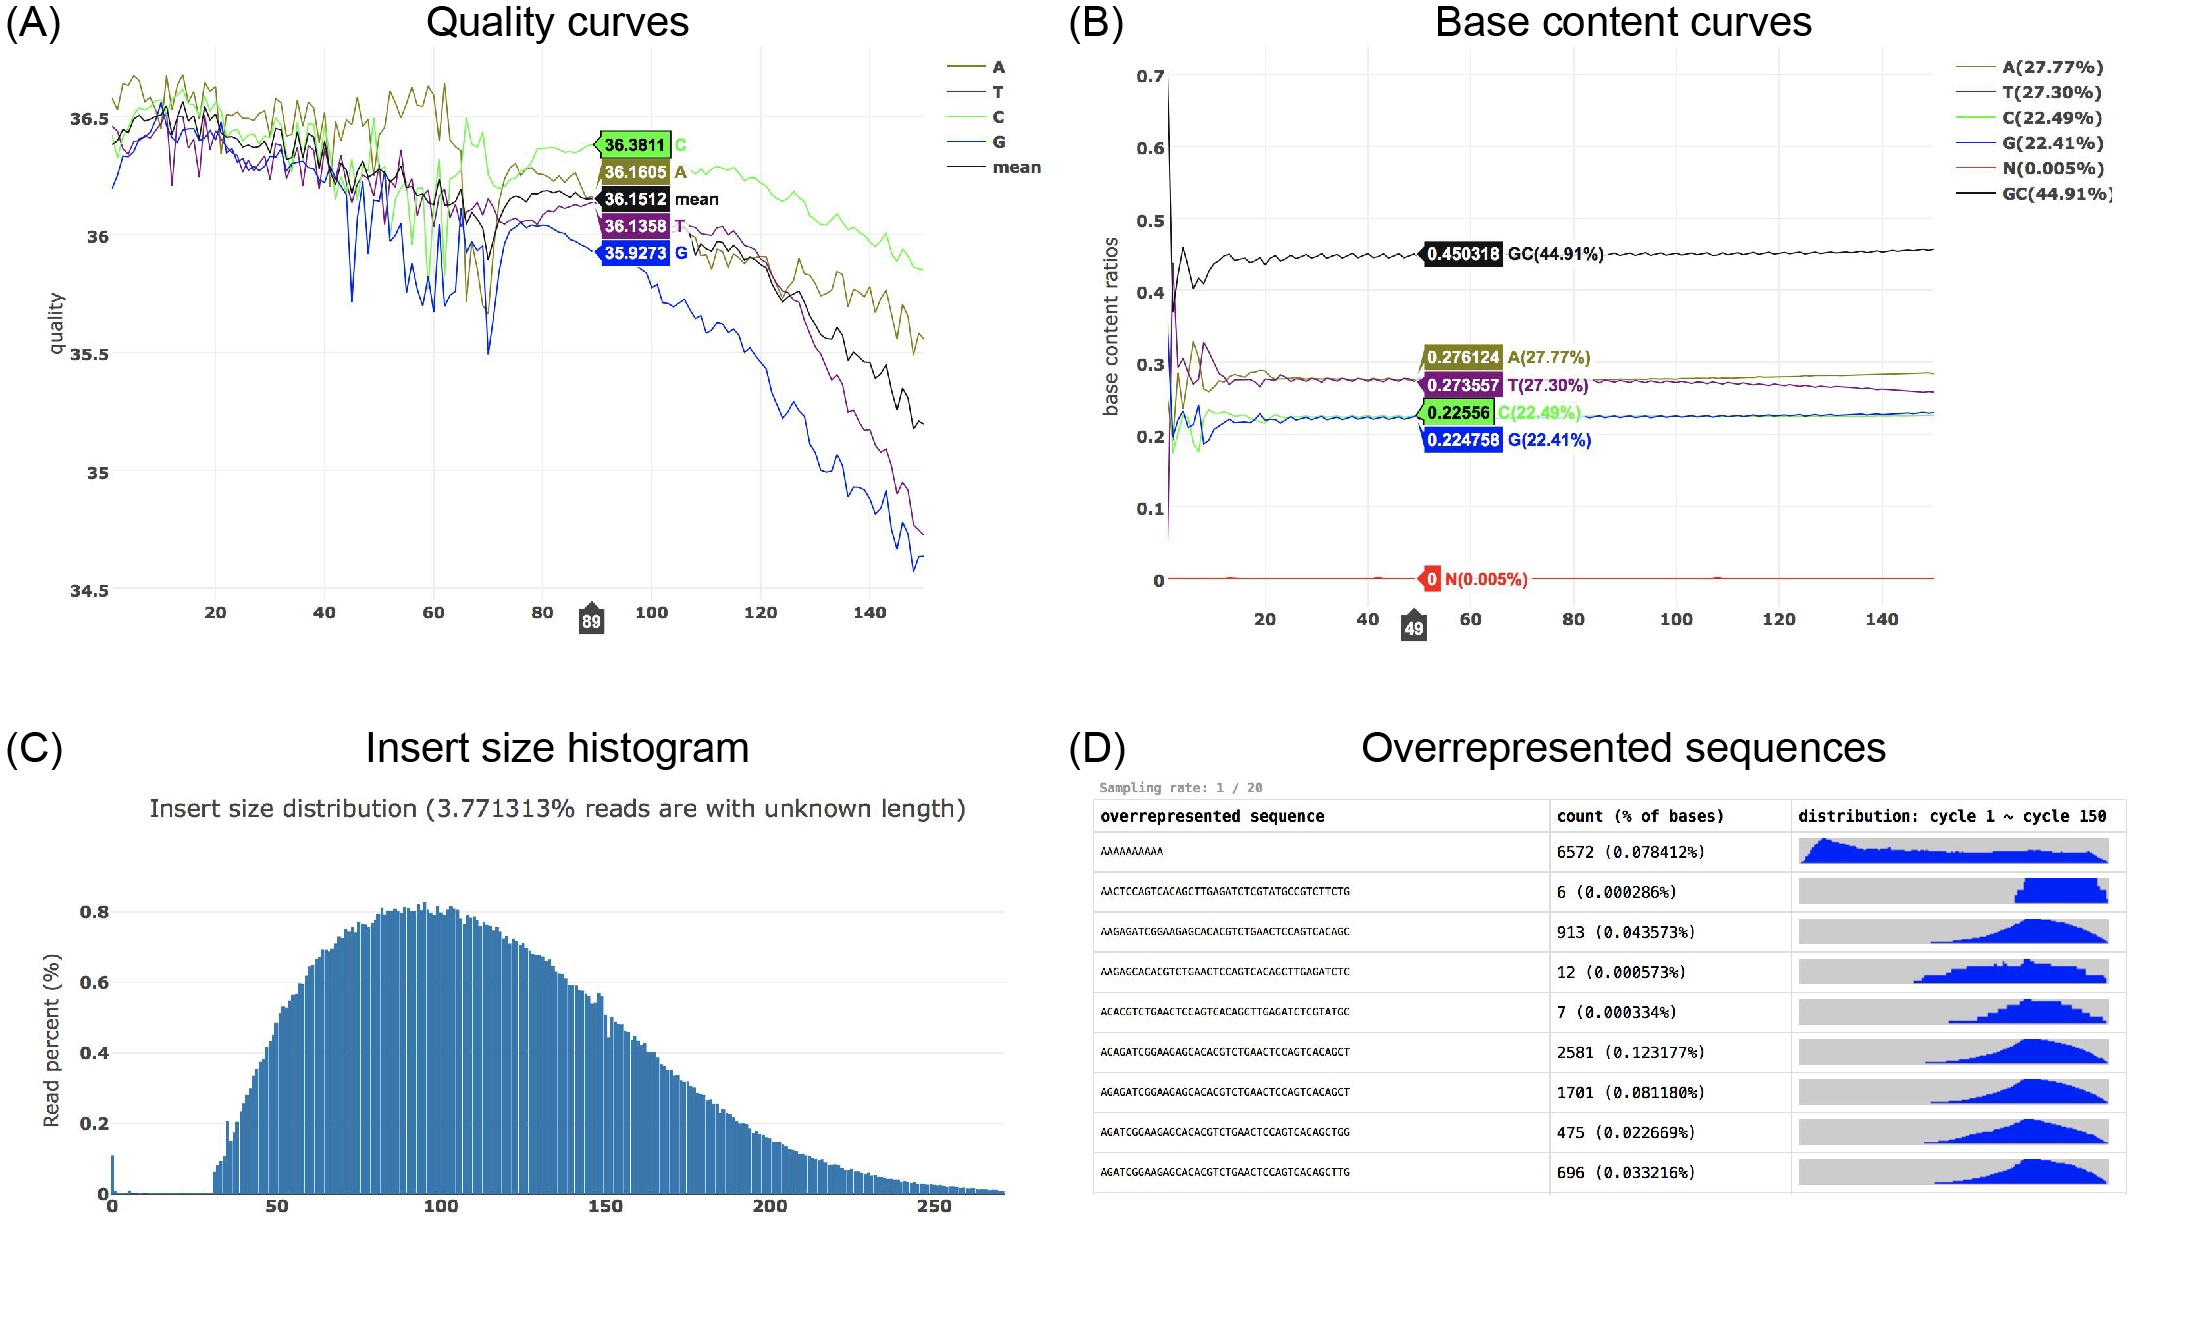
Fastp is a widely adopted tool for FASTQ data preprocessing and quality control. It is ultrafast and versatile and can perform adapter removal, global or quality trimming, read filtering, unique molecular identifier processing, base correction, and many other actions within a single pass of data scanning. Fastp has been reconstructed and upgraded with some new features. Compared to fastp 0.20.0, the new fastp 0.23.2 is even 80% faster.
ImageGP: An easy‐to‐use data visualization web server for scientific researchers
- 21 February 2022
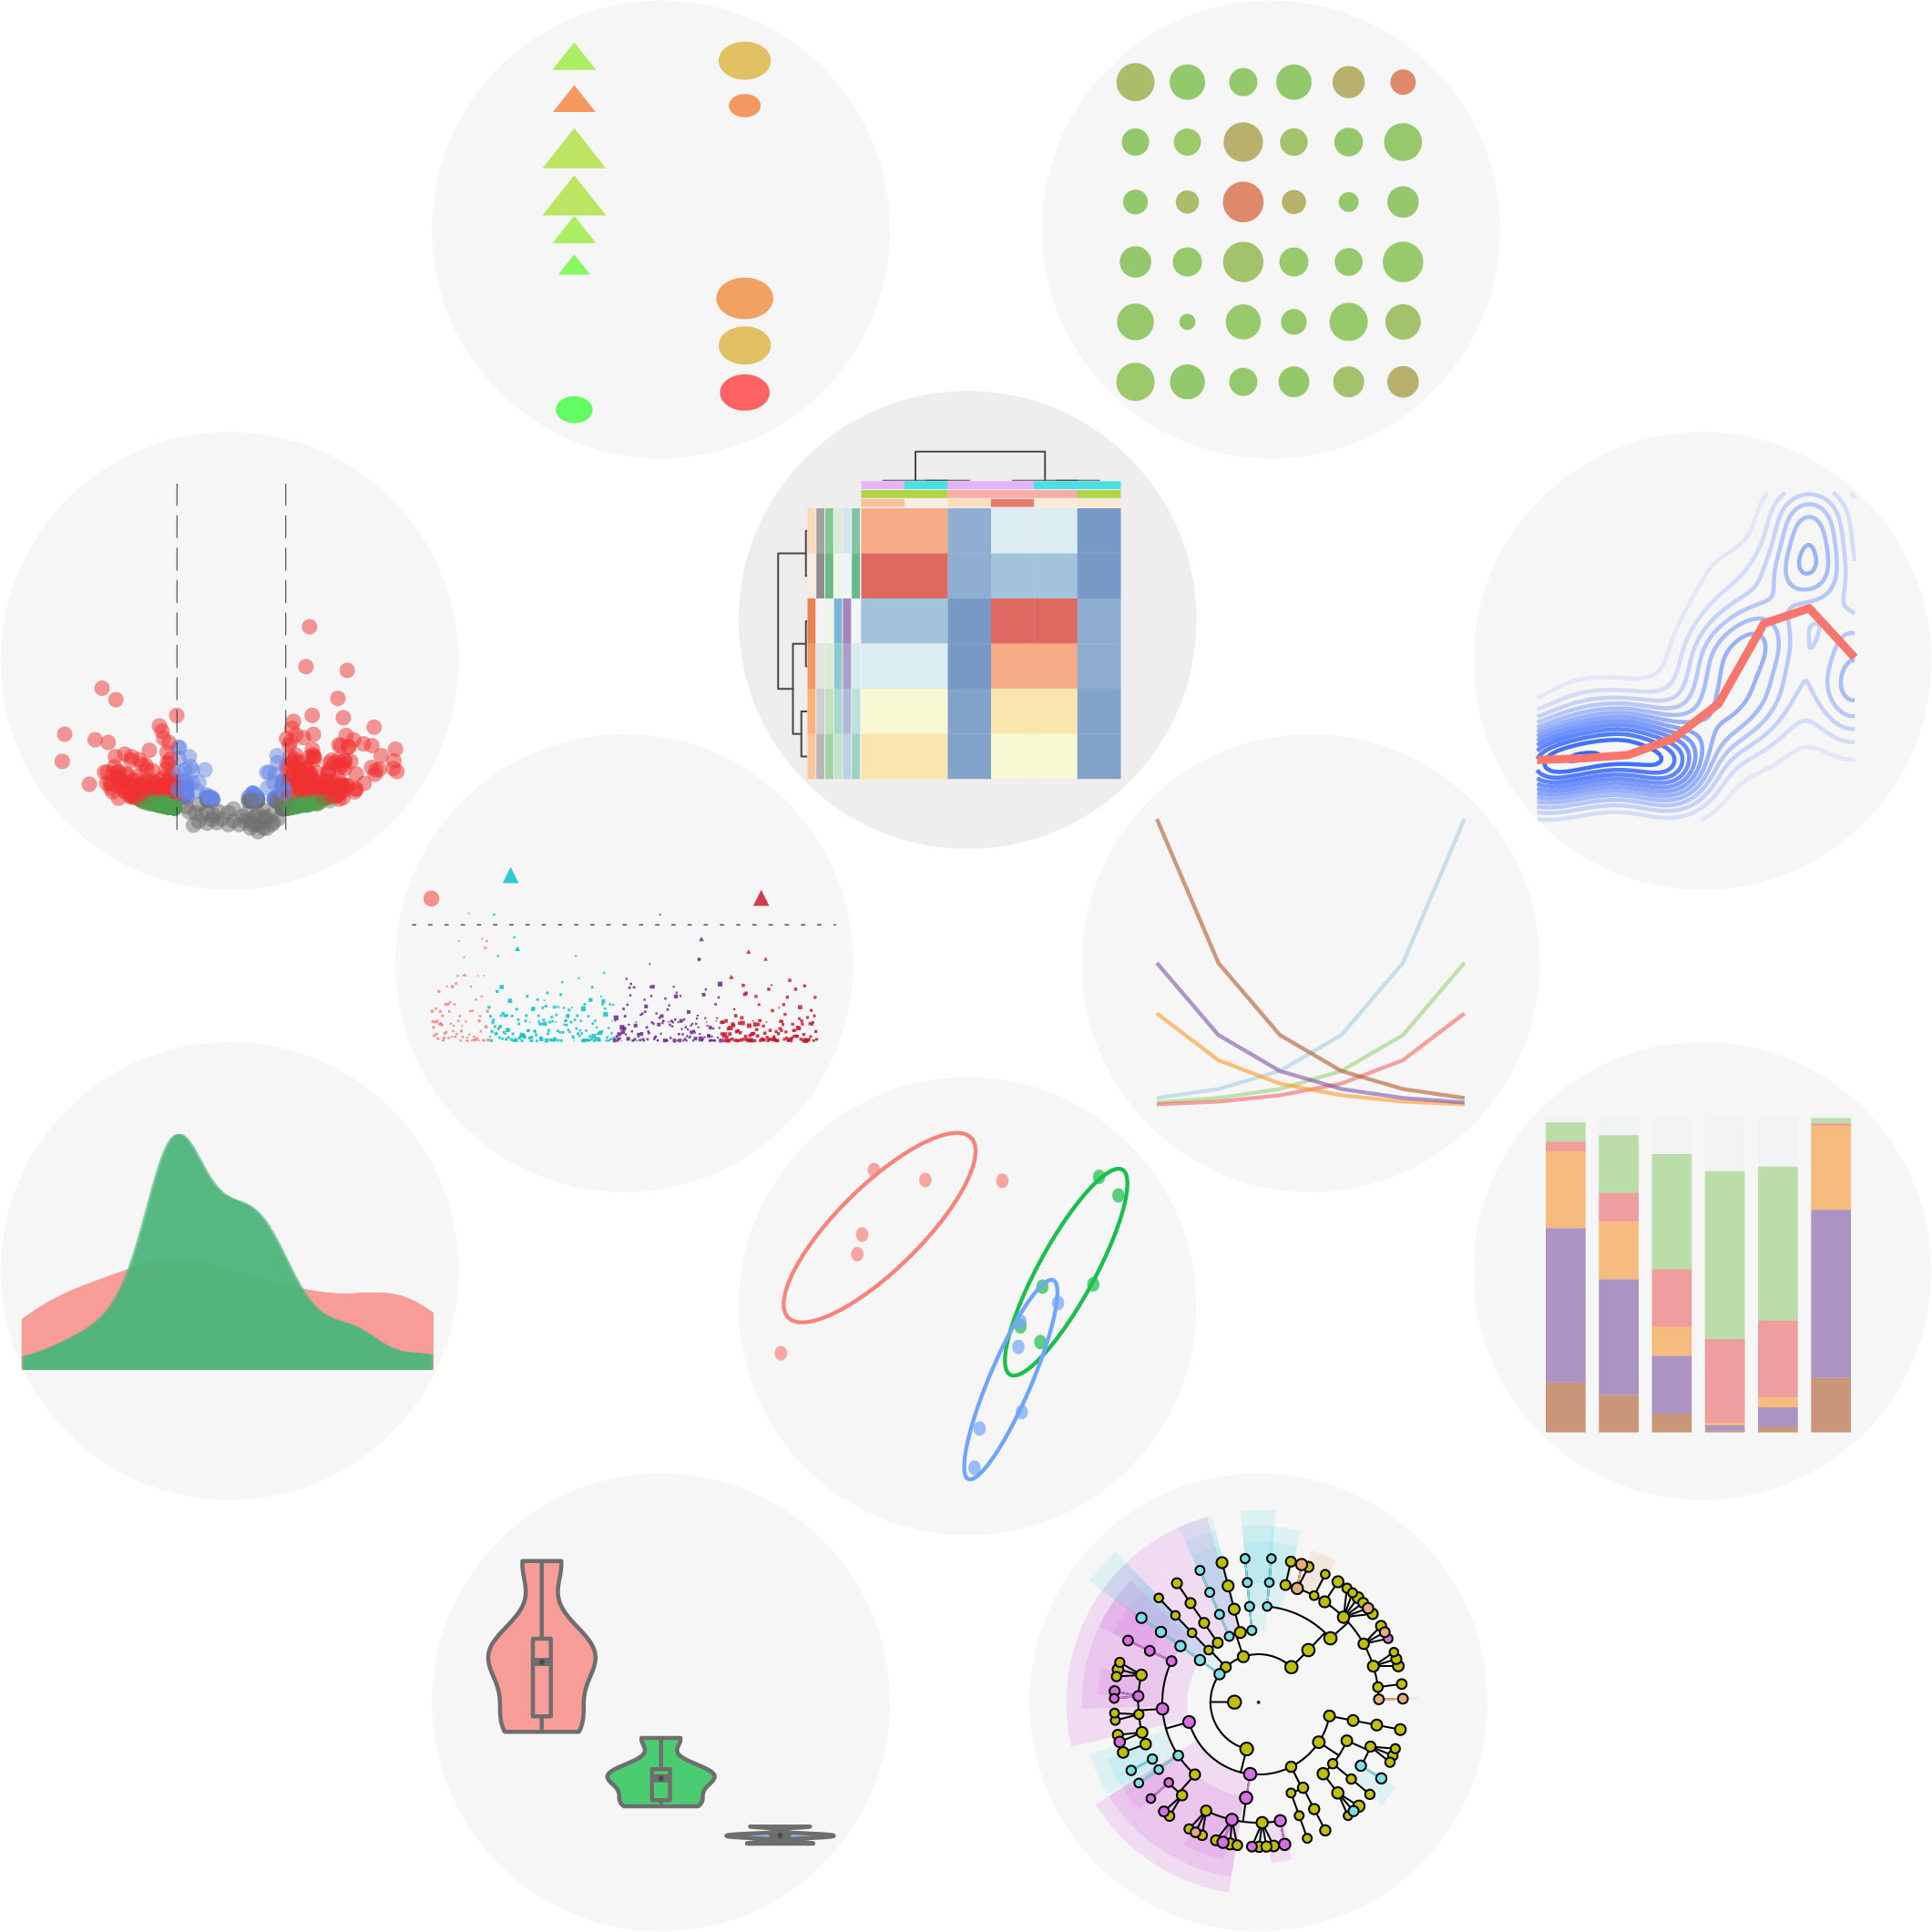
Representative visualization results of ImageGP. ImageGP supports 16 types of images and four types of online analysis with up to 26 parameters for customization. ImageGP also contains specialized plots like volcano plot, functional enrichment plot for most omics-data analysis, and other 4 specialized functions for microbiome analysis. Since 2017, ImageGP has been running for nearly 5 years and serving 336,951 visits from all over the world. Together, ImageGP (http://www.ehbio.com/ImageGP/) is an effective and efficient tool for experimental researchers to comprehensively visualize and interpret data generated from wet-lab and dry-lab.
Ggtree: A serialized data object for visualization of a phylogenetic tree and annotation data
- 28 September 2022
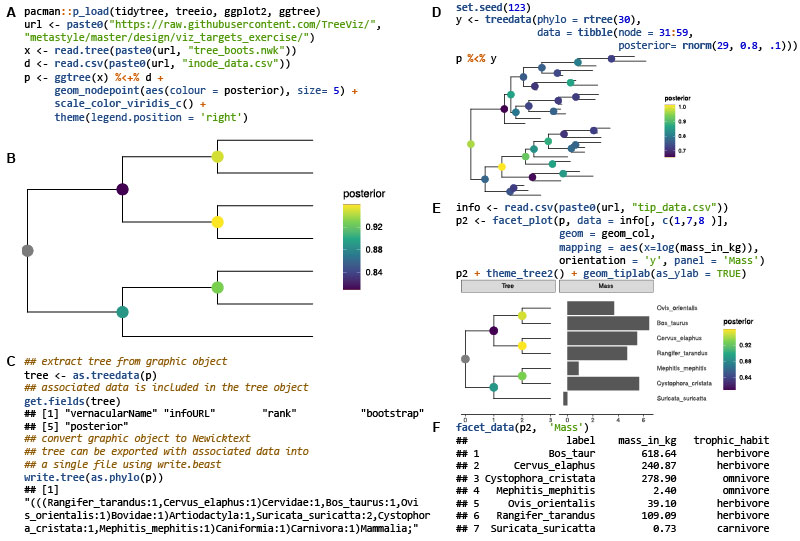
The ggtree object is designed to store phylogenetic tree and associated data, and the object itself is a graphic object that can be rendered as an image file. This work will increase the reproducibility and reusability of phylogenetic data, as well as facilitate integrative and comparative studies.
Using PhyloSuite for molecular phylogeny and tree-based analyses
- 16 February 2023
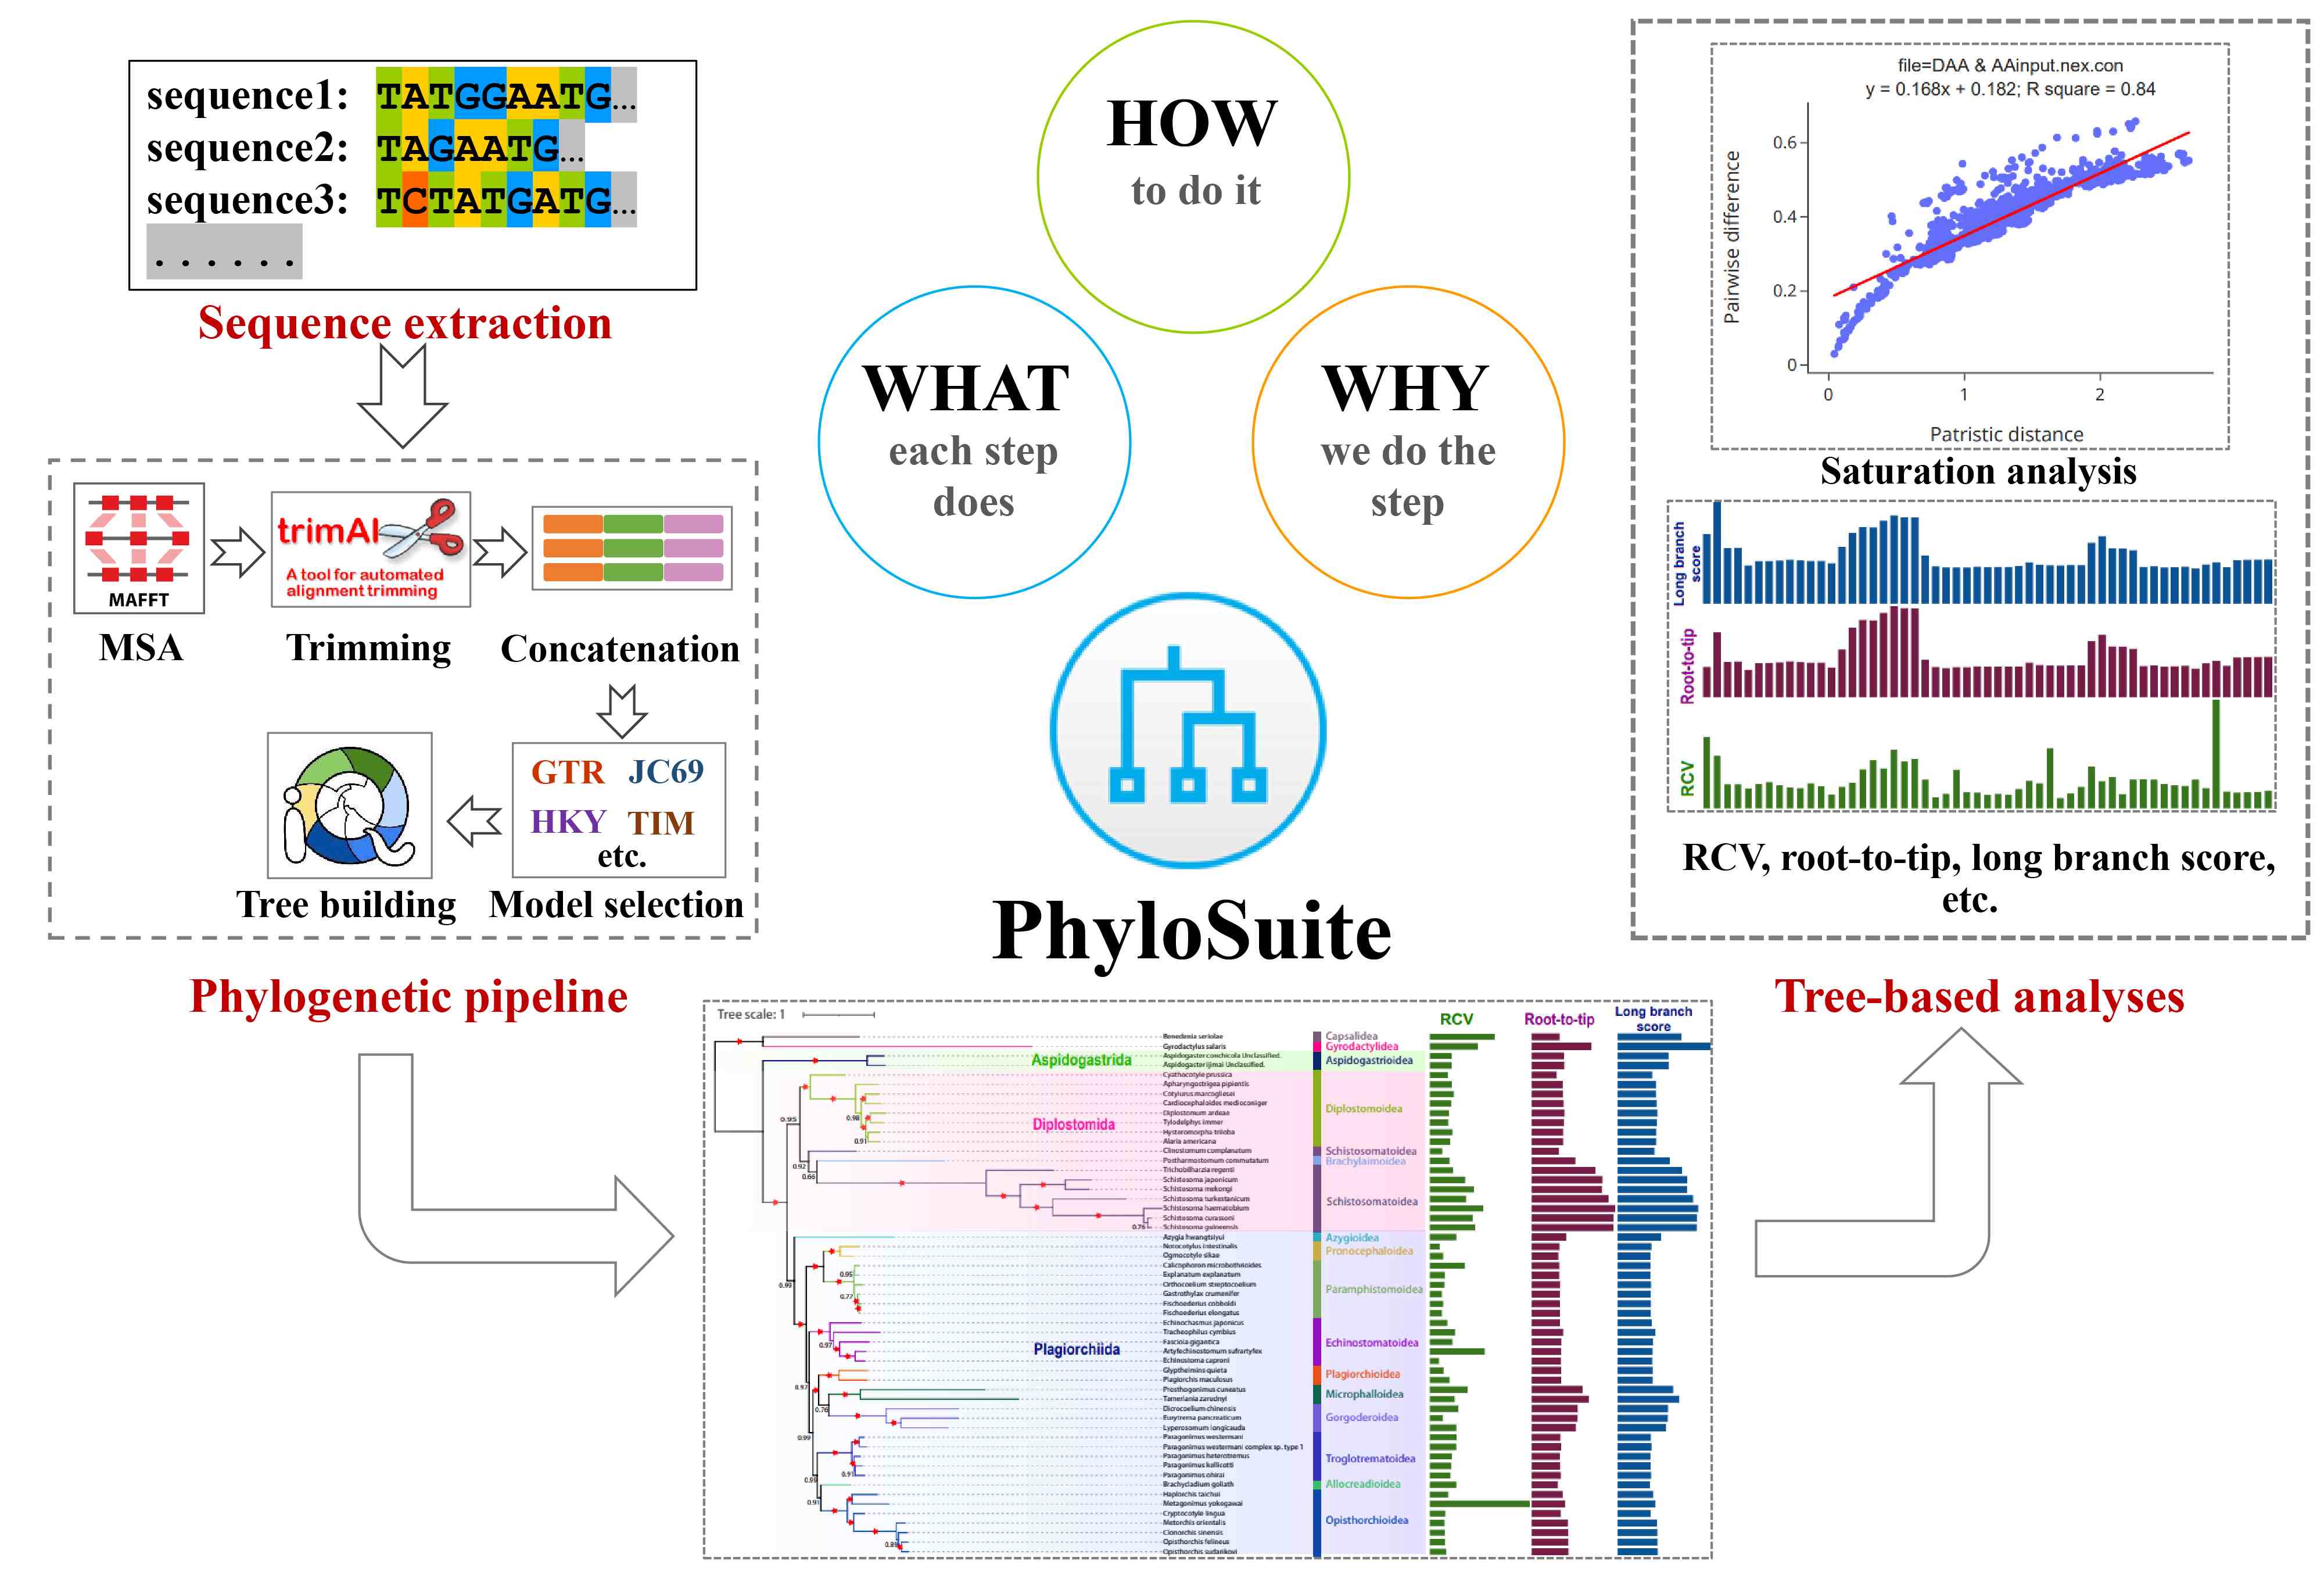
A new release of PhyloSuite, capable of conducting tree-based analyses. Detailed guidelines for each step of phylogenetic and tree-based analyses, following the “What? Why? and How?” structure. This protocol will help beginners learn how to conduct multilocus phylogenetic analyses and help experienced scientists improve their efficiency.